

5 Basics of Molecular Modeling and Molecular Simulation
201
5.2.1.1 Born-Oppenheimer Approximation
Almost all first-principles calculations aimed to solve the many-body Schrödinger
equation have to begin with the Born-Oppenheimer (BO) approximation which
divides the calculations of nuclei and electron interactions into two separate steps.
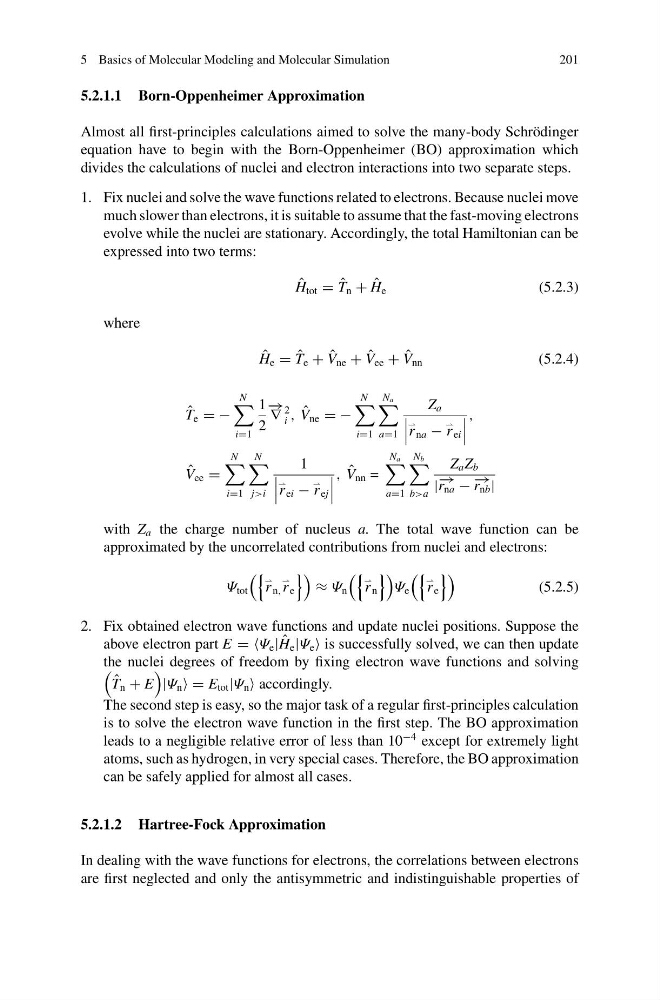
1. Fix nuclei and solve the wave functions related to electrons. Because nuclei move
much slower than electrons, it is suitable to assume that the fast-moving electrons
evolve while the nuclei are stationary. Accordingly, the total Hamiltonian can be
expressed into two terms:
ˆ
H tot = ˆ
T n + ˆ
H e
(5.2.3)
where
ˆ
H e = ˆ
T e + ˆ
V ne + ˆ
V ee + ˆ
V nn
(5.2.4)
ˆ
T e = −
N
i=1
1
2
− →
∇
2
i , ˆ
V ne = −
N
i=1
N a
a=1
Z a
r na −
r ei
,
ˆ
V ee =
N
i=1
N
j>i
1
r ei −
r ej
, ˆ
V nn =
N a
a=1
N b
b>a
Z a Z b
| − →
r na − − →
r nb |
with Z a the charge number of nucleus a. The total wave function can be
approximated by the uncorrelated contributions from nuclei and electrons:
Ψ tot
r n,
r e
≈ Ψ n
r n
Ψ e
r e
(5.2.5)
2. Fix obtained electron wave functions and update nuclei positions. Suppose the
above electron part E = Ψ e | ˆ
H e |Ψ e is successfully solved, we can then update
the nuclei degrees of freedom by fixing electron wave functions and solving
ˆ
T n + E
|Ψ n = E tot |Ψ n accordingly.
The second step is easy, so the major task of a regular first-principles calculation
is to solve the electron wave function in the first step. The BO approximation
leads to a negligible relative error of less than 10
−4 except for extremely light
atoms, such as hydrogen, in very special cases. Therefore, the BO approximation
can be safely applied for almost all cases.
5.2.1.2 Hartree-Fock Approximation
In dealing with the wave functions for electrons, the correlations between electrons
are first neglected and only the antisymmetric and indistinguishable properties of