

130
A. Łachma´ nska et al.
all singly, doubly, triply, etc. substituted configurations. Most commonly, the FCI
expansion is truncated to include only single and double excitations leading to the
CI Singles Doubles (CISD) wave function,
Ψ
CISD
el
= Φ 0 +
occ
i
virt
a
c
a
i a
†
a a i Φ 0 +
occ
i< j
virt
a
ab
i j a
†
a a
†
b a j a i Φ 0 .
(15)
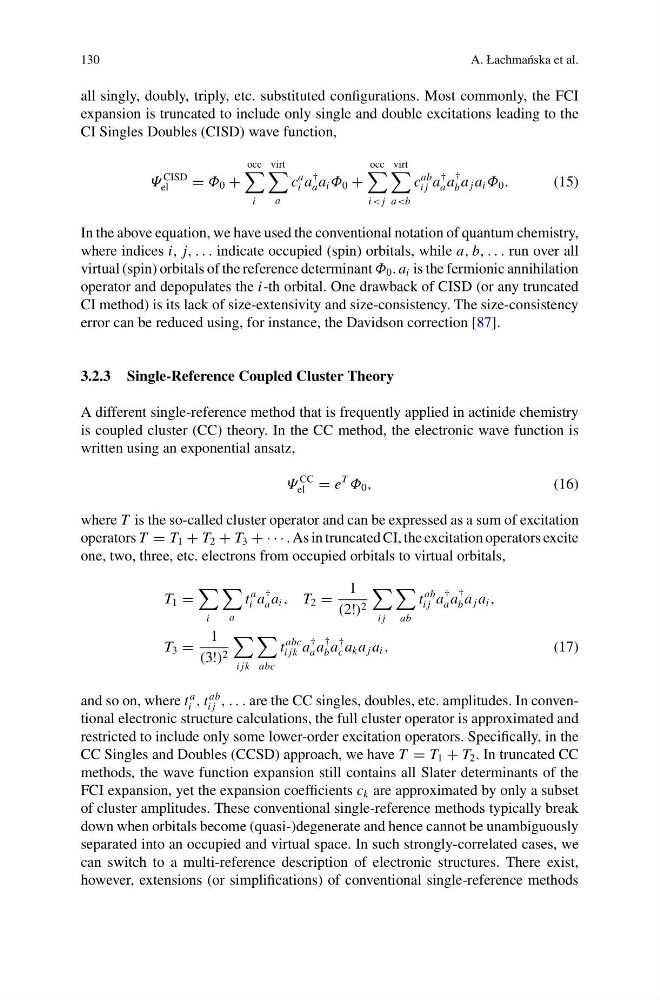
In the above equation, we have used the conventional notation of quantum chemistry,
where indices i, j, . . . indicate occupied (spin) orbitals, while a, b, . . . run over all
virtual (spin) orbitals of the reference determinant Φ 0 . a i is the fermionic annihilation
operator and depopulates the i-th orbital. One drawback of CISD (or any truncated
CI method) is its lack of size-extensivity and size-consistency. The size-consistency
error can be reduced using, for instance, the Davidson correction [87].
3.2.3 Single-Reference Coupled Cluster Theory
A different single-reference method that is frequently applied in actinide chemistry
is coupled cluster (CC) theory. In the CC method, the electronic wave function is
written using an exponential ansatz,
Ψ
CC
el = e
T
Φ 0 ,
(16)
where T is the so-called cluster operator and can be expressed as a sum of excitation
operators T = T 1 + T 2 + T 3 + · · · . As in truncated CI, the excitation operators excite
one, two, three, etc. electrons from occupied orbitals to virtual orbitals,
T 1 =
i
a
t
a
i a
†
a a i , T 2 =
1
(2!) 2
i j
ab
t
ab
i j a
†
a a
†
b a j a i ,
T 3 =
1
(3!) 2
i jk
abc
t
abc
i jk a
†
a a
†
b a
†
c a k a j a i ,
(17)
and so on, where t
a
i , t
ab
i j , . . . are the CC singles, doubles, etc. amplitudes. In conventional electronic structure calculations, the full cluster operator is approximated and
restricted to include only some lower-order excitation operators. Specifically, in the
CC Singles and Doubles (CCSD) approach, we have T = T 1 + T 2 . In truncated CC
methods, the wave function expansion still contains all Slater determinants of the
FCI expansion, yet the expansion coefficients c k are approximated by only a subset
of cluster amplitudes. These conventional single-reference methods typically break
down when orbitals become (quasi-)degenerate and hence cannot be unambiguously
separated into an occupied and virtual space. In such strongly-correlated cases, we
can switch to a multi-reference description of electronic structures. There exist,
however, extensions (or simplifications) of conventional single-reference methods