

26
Chapter 2 Pauling “3-Electron Bonds”, 4-Electron 3-Centre Bonding, and the Need …
differs from those for
B
A
Y
Ψ
and
B
A
Y
Ψ
(whereas they
are the same in the linear combination above). Thus for 3
O , Gould and Linnett
have calculated the “best” linear combination to be
17a
6
5
4
3
2
1
Ψ
Ψ
Ψ
Ψ
Ψ
Ψ
Ψ
028
.
0
124
.
0
390
.
0
351
.
0
best
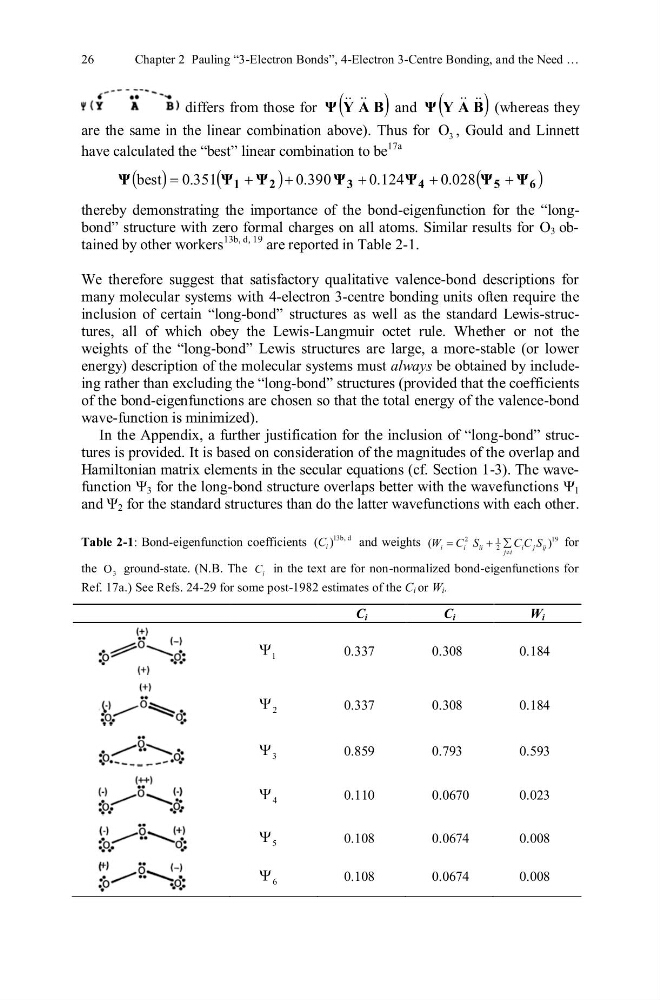
thereby demonstrating the importance of the bond-eigenfunction for the “longbond” structure with zero formal charges on all atoms. Similar results for O 3 obtained by other workers
13b, d, 19 are reported in Table 2-1.
We therefore suggest that satisfactory qualitative valence-bond descriptions for
many molecular systems with 4-electron 3-centre bonding units often require the
inclusion of certain “long-bond” structures as well as the standard Lewis-structures, all of which obey the Lewis-Langmuir octet rule. Whether or not the
weights of the “long-bond” Lewis structures are large, a more-stable (or lower
energy) description of the molecular systems must always be obtained by includeing rather than excluding the “long-bond” structures (provided that the coefficients
of the bond-eigenfunctions are chosen so that the total energy of the valence-bond
wave-function is minimized).
In the Appendix, a further justification for the inclusion of “long-bond” structures is provided. It is based on consideration of the magnitudes of the overlap and
Hamiltonian matrix elements in the secular equations (cf. Section 1-3). The wavefunction Ψ 3 for the long-bond structure overlaps better with the wavefunctions Ψ 1
and Ψ 2 for the standard structures than do the latter wavefunctions with each other.
Table 2-1: Bond-eigenfunction coefficients
13b, d
( )
i
C
and weights
2
19
1
2
(
)
i
i
ii
i j ij
j i
W C S
C C S
for
the 3
O ground-state. (N.B. The i
C in the text are for non-normalized bond-eigenfunctions for
Ref. 17a.) See Refs. 24-29 for some post-1982 estimates of the Ci or Wi.
C i
C i
W i
1
0.337
0.308
0.184
2
0.337
0.308
0.184
3
0.859
0.793
0.593
4
0.110
0.0670
0.023
5
0.108
0.0674
0.008
6
0.108
0.0674
0.008