

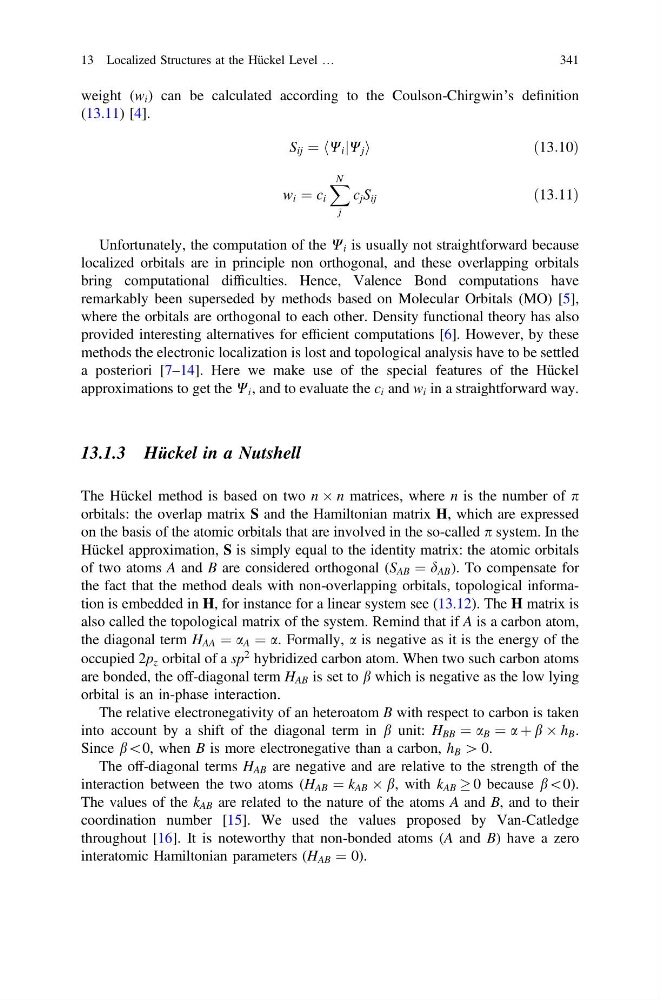
weight (w i ) can be calculated according to the Coulson-Chirgwin’s definition
(13.11) [4].
S ij ¼ hW i jW j i
ð 13:10Þ
w i ¼ c i
X N
j
c j S ij
ð13:11Þ
Unfortunately, the computation of the W i is usually not straightforward because
localized orbitals are in principle non orthogonal, and these overlapping orbitals
bring computational difficulties. Hence, Valence Bond computations have
remarkably been superseded by methods based on Molecular Orbitals (MO) [5],
where the orbitals are orthogonal to each other. Density functional theory has also
provided interesting alternatives for efficient computations [6]. However, by these
methods the electronic localization is lost and topological analysis have to be settled
a posteriori [7–14]. Here we make use of the special features of the Hückel
approximations to get the W i , and to evaluate the c i and w i in a straightforward way.
13.1.3 Hückel in a Nutshell
The Hückel method is based on two n  n matrices, where n is the number of p
orbitals: the overlap matrix S and the Hamiltonian matrix H, which are expressed
on the basis of the atomic orbitals that are involved in the so-called p system. In the
Hückel approximation, S is simply equal to the identity matrix: the atomic orbitals
of two atoms A and B are considered orthogonal (S AB ¼ d AB ). To compensate for
the fact that the method deals with non-overlapping orbitals, topological information is embedded in H, for instance for a linear system see (13.12). The H matrix is
also called the topological matrix of the system. Remind that if A is a carbon atom,
the diagonal term H AA ¼ a A ¼ a. Formally, a is negative as it is the energy of the
occupied 2p z orbital of a sp
2 hybridized carbon atom. When two such carbon atoms
are bonded, the off-diagonal term H AB is set to b which is negative as the low lying
orbital is an in-phase interaction.
The relative electronegativity of an heteroatom B with respect to carbon is taken
into account by a shift of the diagonal term in b unit: H BB ¼ a B ¼ a þ b  h B .
Since b\0, when B is more electronegative than a carbon, h B [ 0.
The off-diagonal terms H AB are negative and are relative to the strength of the
interaction between the two atoms (H AB ¼ k AB Â b, with k AB ! 0 because b\0).
The values of the k AB are related to the nature of the atoms A and B, and to their
coordination number [15]. We used the values proposed by Van-Catledge
throughout [16]. It is noteworthy that non-bonded atoms (A and B) have a zero
interatomic Hamiltonian parameters (H AB ¼ 0).
13 Localized Structures at the Hückel Level …
341