

2.5 Vibrational and Rotational States
53
are the normal coordinates. Note that, as Q = 0 is a minimum of U , ω
2
r ≥ 0 for
all coordinates; i.e., the ω r are all real. The potential U cannot depend on the 6
coordinates describing translations and rotations of the system as a whole; therefore,
6 of the ω r will be equal to 0 (5 in the case R eq corresponds to a linear arrangement
of nuclei). In terms of the normal coordinates ˆ
H vib is given by a sum of equivalent
terms, each one depending on a single Q r
ˆ
H vib = U 0 +
r
ˆ
h harm (Q r )
ˆ
h harm (Q r ) = −
2
2
∂
2
∂ Q 2
r
+
1
2
ω
2
r Q
2
r
(2.121)
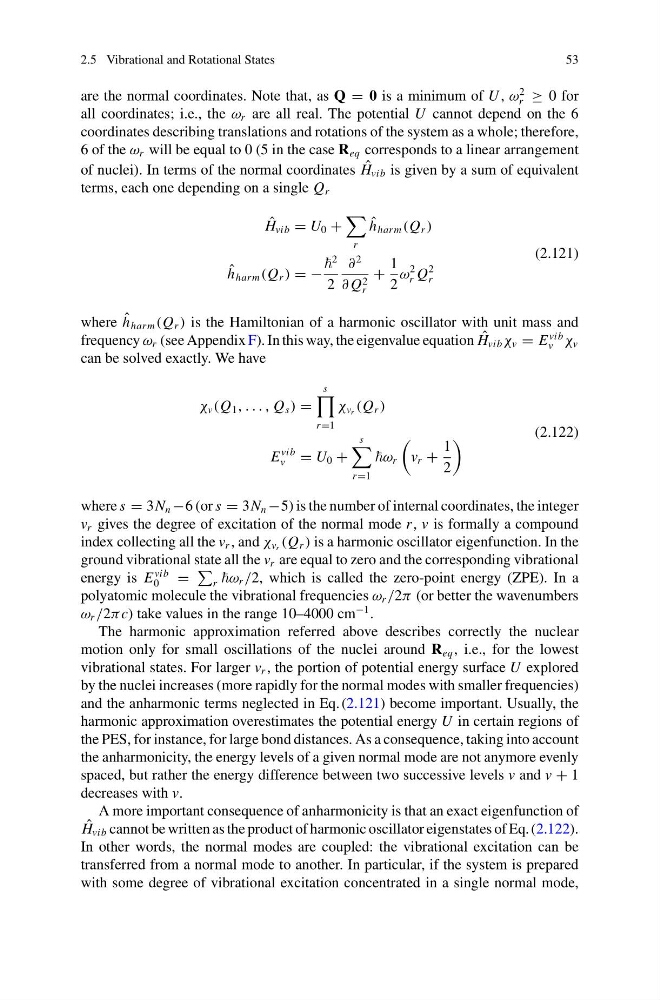
where ˆ
h harm (Q r ) is the Hamiltonian of a harmonic oscillator with unit mass and
frequency ω r (see Appendix F). In this way, the eigenvalue equation ˆ
H vib χ v = E
vib
v χ v
can be solved exactly. We have
χ v (Q 1 , . . . , Q s ) =
s
r =1
χ v r (Q r )
E
vib
v = U 0 +
s
r =1
ω r
v r +
1
2
(2.122)
where s = 3N n −6 (or s = 3N n −5) is the number of internal coordinates, the integer
v r gives the degree of excitation of the normal mode r , v is formally a compound
index collecting all the v r , and χ v r (Q r ) is a harmonic oscillator eigenfunction. In the
ground vibrational state all the v r are equal to zero and the corresponding vibrational
energy is E
vib
0
=
r ω r /2, which is called the zero-point energy (ZPE). In a
polyatomic molecule the vibrational frequencies ω r /2π (or better the wavenumbers
ω r /2π c) take values in the range 10–4000 cm
−1 .
The harmonic approximation referred above describes correctly the nuclear
motion only for small oscillations of the nuclei around R eq , i.e., for the lowest
vibrational states. For larger v r , the portion of potential energy surface U explored
by the nuclei increases (more rapidly for the normal modes with smaller frequencies)
and the anharmonic terms neglected in Eq. (2.121) become important. Usually, the
harmonic approximation overestimates the potential energy U in certain regions of
the PES, for instance, for large bond distances. As a consequence, taking into account
the anharmonicity, the energy levels of a given normal mode are not anymore evenly
spaced, but rather the energy difference between two successive levels v and v + 1
decreases with v.
A more important consequence of anharmonicity is that an exact eigenfunction of
ˆ
H vib cannot be written as the product of harmonic oscillator eigenstates of Eq. (2.122).
In other words, the normal modes are coupled: the vibrational excitation can be
transferred from a normal mode to another. In particular, if the system is prepared
with some degree of vibrational excitation concentrated in a single normal mode,