

5.1 Toward More Complex Systems
159
from the usual expansion in partial waves proceeds then through the steps already
singled out in the previous chapter for the atom–diatom systems. This implies the
carrying out of the propagation of the solution from the first sector to the last one,
matching the calculated values with the asymptotic ones for all the necessary boundary conditions, working out of this comparison the value of the elements of the S
matrix, performing the appropriate statistical averaging of the computed quantities
to the end of calculating the corresponding observables. These steps, though conceptually identical to those of the atom–diatom case, become increasingly more difficult
for heavier systems, more complex electronic structures and a larger coupling of the
related calculations. For this purpose, large use of the package MOLSCATT [103]
based on an expansion of the potential in terms of spherical harmonics, is made for
nonreactive systems.
For reactive processes, it is usually preferred to adopt the hyperspherical formalism that defines the reaction coordinate as the hyperradius (regardless of the
arrangement number and type of the atoms composing the molecular subsystem).
However, the increasing difficulty of dealing with the fixed ρ surface functions (the
eigenfunctions of the 3N-4 hyperangles) for N larger than 4, has till now made
the generalized use of these coordinates computationally impractical for scattering
calculations even if formally interesting.
For four-atom (diatom–diatom in our case) systems one can reduce the complexity of the problem using the Quantum-Classical (QC) Coupled-Channel method
(see Refs. [104–106] for a more extended discussion) in which molecular vibrations are treated quantum-mechanically by integrating the related time-dependent
Schrödinger equations for the N 2 and the O 2 molecules. On the contrary, translational and rotational degrees of freedom are treated classically by integrating the
related classical Hamilton equations. The two subsystems, and the corresponding
equations of motion, are dynamically coupled through the definition and calculation
of a time-dependent “effective” Hamiltonian, of the Ehrenfest type, defined as the
expectation value of the intermolecular interaction potential over (r a , r b , t)
H eff =< <(r a , r b , t) | V inter (R(t)) | (r a , r b , t) >
(5.11)
where V inter (R(t)) is the intermolecular interaction potential evaluated at each time
step of the classical “mean” trajectory R(t).
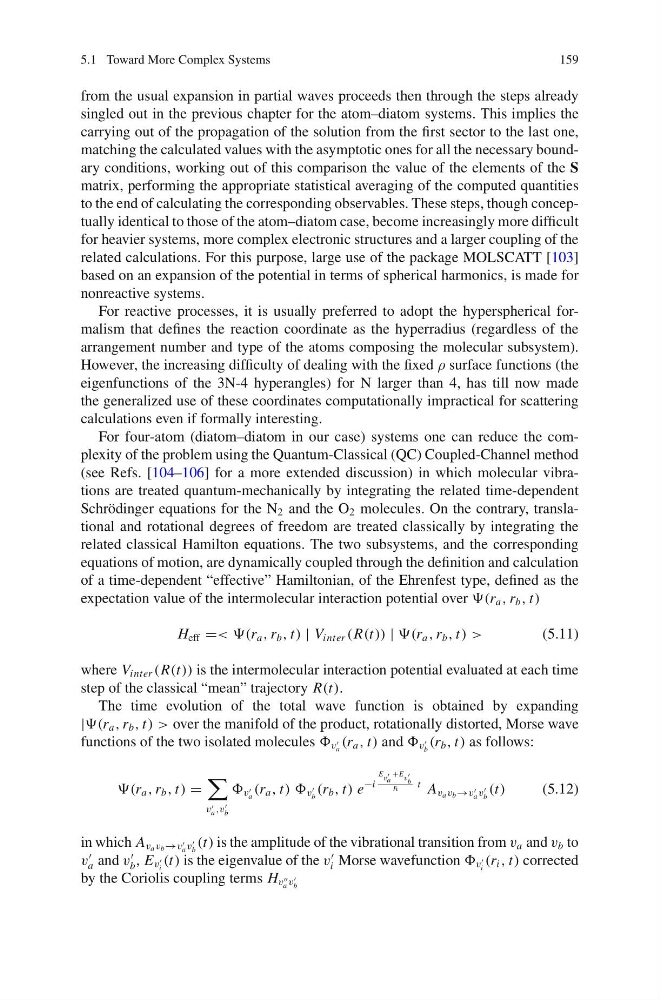
The time evolution of the total wave function is obtained by expanding
|(r a , r b , t) > over the manifold of the product, rotationally distorted, Morse wave
functions of the two isolated molecules v
a
(r a , t) and v
b
(r b , t) as follows:
(r a , r b , t) =
v
a ,v
b
v
a
(r a , t) ) v
b
(r b , t) e
−i
E v
a
+E v
b
t A v a v b →v
a v
b
(t)
(5.12)
in which A v a v b →v
a v
b
(t) is the amplitude of the vibrational transition from v a and v b to
v
a and v
b , E v
i
(t) is the eigenvalue of the v
i Morse wavefunction v
i
(r i , t) corrected
by the Coriolis coupling terms H v
a v
b