

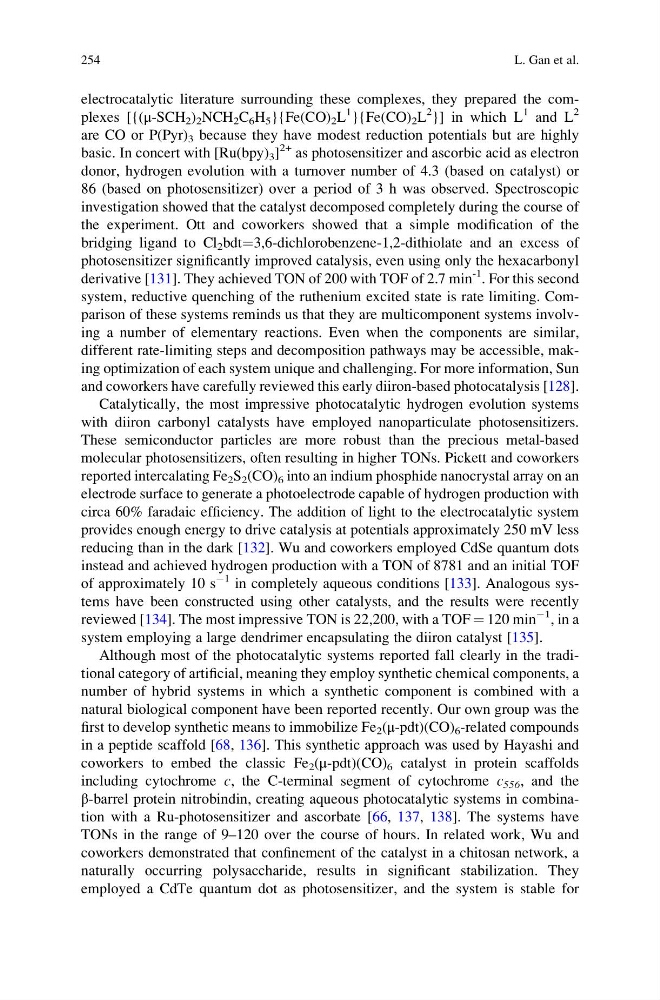
electrocatalytic literature surrounding these complexes, they prepared the complexes [{(μ-SCH 2 ) 2 NCH 2 C 6 H 5 }{Fe(CO) 2 L
1 }{Fe(CO) 2 L
2 }] in which L
1 and L
2
are CO or P(Pyr) 3 because they have modest reduction potentials but are highly
basic. In concert with [Ru(bpy) 3 ]
2+ as photosensitizer and ascorbic acid as electron
donor, hydrogen evolution with a turnover number of 4.3 (based on catalyst) or
86 (based on photosensitizer) over a period of 3 h was observed. Spectroscopic
investigation showed that the catalyst decomposed completely during the course of
the experiment. Ott and coworkers showed that a simple modification of the
bridging ligand to Cl 2 bdt¼3,6-dichlorobenzene-1,2-dithiolate and an excess of
photosensitizer significantly improved catalysis, even using only the hexacarbonyl
derivative [131]. They achieved TON of 200 with TOF of 2.7 min
-1 . For this second
system, reductive quenching of the ruthenium excited state is rate limiting. Comparison of these systems reminds us that they are multicomponent systems involving a number of elementary reactions. Even when the components are similar,
different rate-limiting steps and decomposition pathways may be accessible, making optimization of each system unique and challenging. For more information, Sun
and coworkers have carefully reviewed this early diiron-based photocatalysis [128].
Catalytically, the most impressive photocatalytic hydrogen evolution systems
with diiron carbonyl catalysts have employed nanoparticulate photosensitizers.
These semiconductor particles are more robust than the precious metal-based
molecular photosensitizers, often resulting in higher TONs. Pickett and coworkers
reported intercalating Fe 2 S 2 (CO) 6 into an indium phosphide nanocrystal array on an
electrode surface to generate a photoelectrode capable of hydrogen production with
circa 60% faradaic efficiency. The addition of light to the electrocatalytic system
provides enough energy to drive catalysis at potentials approximately 250 mV less
reducing than in the dark [132]. Wu and coworkers employed CdSe quantum dots
instead and achieved hydrogen production with a TON of 8781 and an initial TOF
of approximately 10 s
À1 in completely aqueous conditions [133]. Analogous systems have been constructed using other catalysts, and the results were recently
reviewed [134]. The most impressive TON is 22,200, with a TOF ¼ 120 min
À1 , in a
system employing a large dendrimer encapsulating the diiron catalyst [135].
Although most of the photocatalytic systems reported fall clearly in the traditional category of artificial, meaning they employ synthetic chemical components, a
number of hybrid systems in which a synthetic component is combined with a
natural biological component have been reported recently. Our own group was the
first to develop synthetic means to immobilize Fe 2 (μ-pdt)(CO) 6 -related compounds
in a peptide scaffold [68, 136]. This synthetic approach was used by Hayashi and
coworkers to embed the classic Fe 2 (μ-pdt)(CO) 6 catalyst in protein scaffolds
including cytochrome c, the C-terminal segment of cytochrome c 556 , and the
β-barrel protein nitrobindin, creating aqueous photocatalytic systems in combination with a Ru-photosensitizer and ascorbate [66, 137, 138]. The systems have
TONs in the range of 9–120 over the course of hours. In related work, Wu and
coworkers demonstrated that confinement of the catalyst in a chitosan network, a
naturally occurring polysaccharide, results in significant stabilization. They
employed a CdTe quantum dot as photosensitizer, and the system is stable for
254
L. Gan et al.