

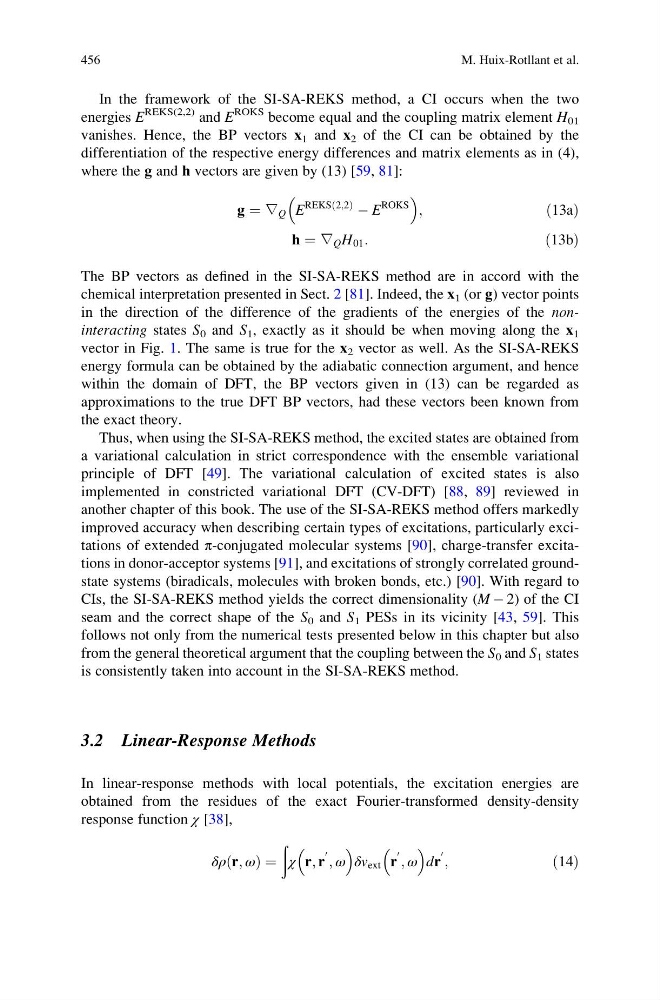
In the framework of the SI-SA-REKS method, a CI occurs when the two
energies E
REKS(2,2) and E
ROKS become equal and the coupling matrix element H 01
vanishes. Hence, the BP vectors x 1 and x 2 of the CI can be obtained by the
differentiation of the respective energy differences and matrix elements as in (4),
where the g and h vectors are given by (13) [59, 81]:
g ¼ ∇ Q E
REKS 2;2
ð Þ
À E
ROKS
;
ð13aÞ
h ¼ ∇ Q H 01 :
ð13bÞ
The BP vectors as defined in the SI-SA-REKS method are in accord with the
chemical interpretation presented in Sect. 2 [81]. Indeed, the x 1 (or g) vector points
in the direction of the difference of the gradients of the energies of the noninteracting states S 0 and S 1 , exactly as it should be when moving along the x 1
vector in Fig. 1. The same is true for the x 2 vector as well. As the SI-SA-REKS
energy formula can be obtained by the adiabatic connection argument, and hence
within the domain of DFT, the BP vectors given in (13) can be regarded as
approximations to the true DFT BP vectors, had these vectors been known from
the exact theory.
Thus, when using the SI-SA-REKS method, the excited states are obtained from
a variational calculation in strict correspondence with the ensemble variational
principle of DFT [49]. The variational calculation of excited states is also
implemented in constricted variational DFT (CV-DFT) [88, 89] reviewed in
another chapter of this book. The use of the SI-SA-REKS method offers markedly
improved accuracy when describing certain types of excitations, particularly excitations of extended π-conjugated molecular systems [90], charge-transfer excitations in donor-acceptor systems [91], and excitations of strongly correlated groundstate systems (biradicals, molecules with broken bonds, etc.) [90]. With regard to
CIs, the SI-SA-REKS method yields the correct dimensionality (M À 2) of the CI
seam and the correct shape of the S 0 and S 1 PESs in its vicinity [43, 59]. This
follows not only from the numerical tests presented below in this chapter but also
from the general theoretical argument that the coupling between the S 0 and S 1 states
is consistently taken into account in the SI-SA-REKS method.
3.2 Linear-Response Methods
In linear-response methods with local potentials, the excitation energies are
obtained from the residues of the exact Fourier-transformed density-density
response function χ [38],
δρ r; ω
ð
Þ ¼
ð
χ r; r
0 ; ω
δv ext r
0 ; ω
dr
0 ;
ð14Þ
456
M. Huix-Rotllant et al.