

properties has a decisive role. This topic is in details discussed in many Refs. (see,
for example, [13, 14, 51–58, 60–63, 97]).
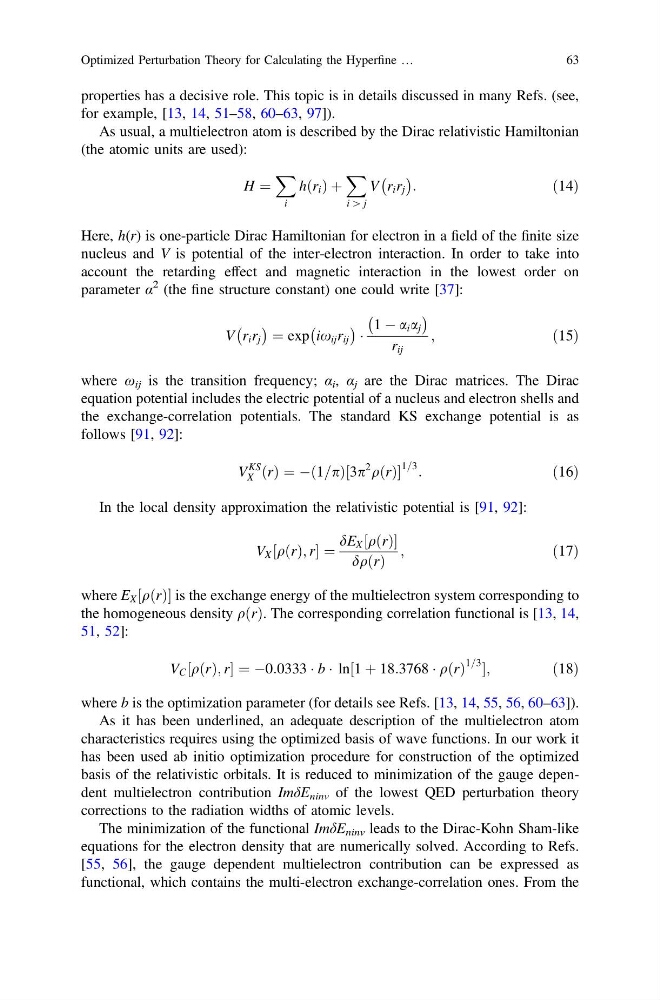
As usual, a multielectron atom is described by the Dirac relativistic Hamiltonian
(the atomic units are used):
H ¼
X
i
hðr i Þ þ
X
i [ j
V r i r j
À Á :
ð14Þ
Here, h(r) is one-particle Dirac Hamiltonian for electron in a field of the finite size
nucleus and V is potential of the inter-electron interaction. In order to take into
account the retarding effect and magnetic interaction in the lowest order on
parameter α
2 (the fine structure constant) one could write [37]:
V r i r j
À Á ¼ exp ix ij r ij
À
Á Á
1 À a i a j
À
Á
r ij
;
ð15Þ
where ω ij is the transition frequency; α i , α j are the Dirac matrices. The Dirac
equation potential includes the electric potential of a nucleus and electron shells and
the exchange-correlation potentials. The standard KS exchange potential is as
follows [91, 92]:
V
KS
X ðrÞ ¼ Àð1=pÞ½3p
2
qðrÞ
1=3 :
ð16Þ
In the local density approximation the relativistic potential is [91, 92]:
V X ½qðrÞ; r ¼
dE X ½qðrÞ
dqðrÞ
;
ð17Þ
where E X ½qðrÞ is the exchange energy of the multielectron system corresponding to
the homogeneous density qðrÞ. The corresponding correlation functional is [13, 14,
51, 52]:
V C ½qðrÞ; r ¼ À0:0333 Á b Á ln½1 þ 18:3768 Á qðrÞ
1=3 ;
ð18Þ
where b is the optimization parameter (for details see Refs. [13, 14, 55, 56, 60–63]).
As it has been underlined, an adequate description of the multielectron atom
characteristics requires using the optimized basis of wave functions. In our work it
has been used ab initio optimization procedure for construction of the optimized
basis of the relativistic orbitals. It is reduced to minimization of the gauge dependent multielectron contribution ImδE ninv of the lowest QED perturbation theory
corrections to the radiation widths of atomic levels.
The minimization of the functional ImδE ninv leads to the Dirac-Kohn Sham-like
equations for the electron density that are numerically solved. According to Refs.
[55, 56], the gauge dependent multielectron contribution can be expressed as
functional, which contains the multi-electron exchange-correlation ones. From the
Optimized Perturbation Theory for Calculating the Hyperfine …
63