

7 First-Principles Investigations of Electronically …
173
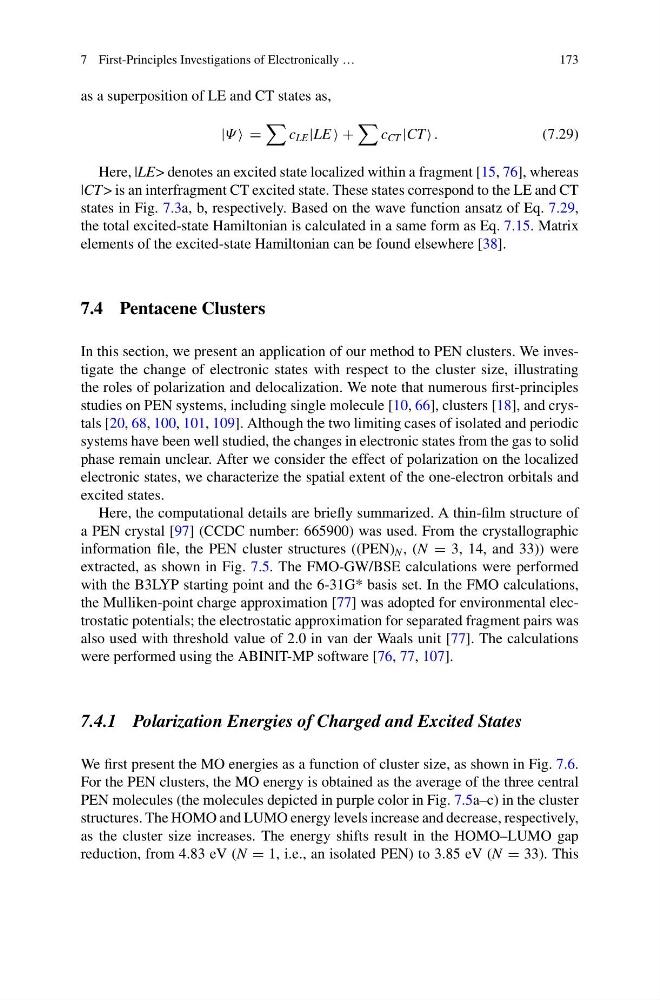
as a superposition of LE and CT states as,
|Ψ =
c LE |LE +
c CT |CT .
(7.29)
Here, |LE> denotes an excited state localized within a fragment [15, 76], whereas
|CT > is an interfragment CT excited state. These states correspond to the LE and CT
states in Fig. 7.3a, b, respectively. Based on the wave function ansatz of Eq. 7.29,
the total excited-state Hamiltonian is calculated in a same form as Eq. 7.15. Matrix
elements of the excited-state Hamiltonian can be found elsewhere [38].
7.4 Pentacene Clusters
In this section, we present an application of our method to PEN clusters. We investigate the change of electronic states with respect to the cluster size, illustrating
the roles of polarization and delocalization. We note that numerous first-principles
studies on PEN systems, including single molecule [10, 66], clusters [18], and crystals [20, 68, 100, 101, 109]. Although the two limiting cases of isolated and periodic
systems have been well studied, the changes in electronic states from the gas to solid
phase remain unclear. After we consider the effect of polarization on the localized
electronic states, we characterize the spatial extent of the one-electron orbitals and
excited states.
Here, the computational details are briefly summarized. A thin-film structure of
a PEN crystal [97] (CCDC number: 665900) was used. From the crystallographic
information file, the PEN cluster structures ((PEN) N , (N = 3, 14, and 33)) were
extracted, as shown in Fig. 7.5. The FMO-GW/BSE calculations were performed
with the B3LYP starting point and the 6-31G* basis set. In the FMO calculations,
the Mulliken-point charge approximation [77] was adopted for environmental electrostatic potentials; the electrostatic approximation for separated fragment pairs was
also used with threshold value of 2.0 in van der Waals unit [77]. The calculations
were performed using the ABINIT-MP software [76, 77, 107].
7.4.1 Polarization Energies of Charged and Excited States
We first present the MO energies as a function of cluster size, as shown in Fig. 7.6.
For the PEN clusters, the MO energy is obtained as the average of the three central
PEN molecules (the molecules depicted in purple color in Fig. 7.5a–c) in the cluster
structures. The HOMO and LUMO energy levels increase and decrease, respectively,
as the cluster size increases. The energy shifts result in the HOMO–LUMO gap
reduction, from 4.83 eV (N = 1, i.e., an isolated PEN) to 3.85 eV (N = 33). This