

4 Nanoscale First-Principles Electronic Structure Simulations of Materials. . .
105
Energy (eV)
-6.0 -5.0 -4.0 -3.0 -2.0 -1.0 0.0 1.0 2.0 3.0
DOS (states·eV
−1
)
DOS (arb. unit)
(a)
(b)
H−2
H−1
H
H−4
H−3
L
L+1
Exp.
L+2
-4.0
-2.0
0.0
2.0
4.0
X
Y
Energy (eV)
L
Z N
MR
H−2
H−1
H
H−4
H−3
L
L+1
L+2
W H
W L
E g
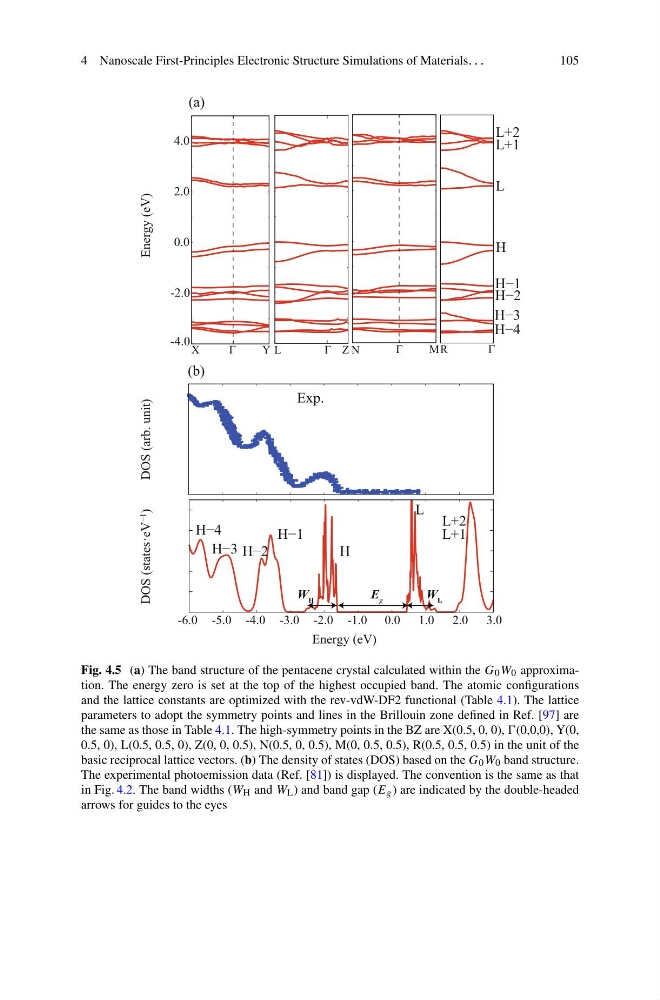
Fig. 4.5 (a) The band structure of the pentacene crystal calculated within the G 0 W 0 approximation. The energy zero is set at the top of the highest occupied band. The atomic configurations
and the lattice constants are optimized with the rev-vdW-DF2 functional (Table 4.1). The lattice
parameters to adopt the symmetry points and lines in the Brillouin zone defined in Ref. [97] are
the same as those in Table 4.1. The high-symmetry points in the BZ are X(0.5, 0, 0), (0,0,0), Y(0,
0.5, 0), L(0.5, 0.5, 0), Z(0, 0, 0.5), N(0.5, 0, 0.5), M(0, 0.5, 0.5), R(0.5, 0.5, 0.5) in the unit of the
basic reciprocal lattice vectors. (b) The density of states (DOS) based on the G 0 W 0 band structure.
The experimental photoemission data (Ref. [81]) is displayed. The convention is the same as that
in Fig. 4.2. The band widths (W H and W L ) and band gap (E g ) are indicated by the double-headed
arrows for guides to the eyes