

162
5 Vibrational Up-Pumping in Polymorphic Materials
Condensed Matter Calculations. All condensed matter vibrational calculations were
based on experimental crystal structures. Structures for δ-HMX (OCHTET03) and
β-FOX-7 (SEDTUQ06) were sourced from the Cambridge Crystallographic Data
Centre (CCDC). The input structure for γ-FOX-7 was taken from Crawford et al.
[25]. Data for β-HMX and α-FOX-7 were taken from Chap. 4. All structures were
optimised using plane-wave DFT (pw-DFT) as implemented in CASTEP v16 [33].
The GGA functional of Perdew-Burke-Ernzerhof [34] (PBE) was used along with
Grimme’s D2 dispersion correction scheme [35], PBE-D2. This scheme has previously been demonstrated to perform well for these materials [21, 36]. The electronic wavefunction was expanded in plane waves to a kinetic energy cut-off of
1800 eV (α-FOX-7), 1300 eV (β-HMX, δ-HMX and β-FOX-7) and 950 eV (γFOX-7). All forces were converged to <5 × 10
−4 eV/Å and stresses to <5 × 10
−4
GPa. γ-FOX-7 was converged to less stringent parameters, forces <1 × 10
−3 eV/
Å and stresses to <1 × 10
−3 GPA. The energy change per atom was accepted after
convergence <5 × 10
−9 eV/atom for all cases except γ-FOX-7, for which a convergence was accepted <1 × 10
−8 eV/atom. The electronic structure was sampled
on a k-point Monkhorst-Pack [37] (MP) grid with spacing no more than 0.05 Å
−1 .
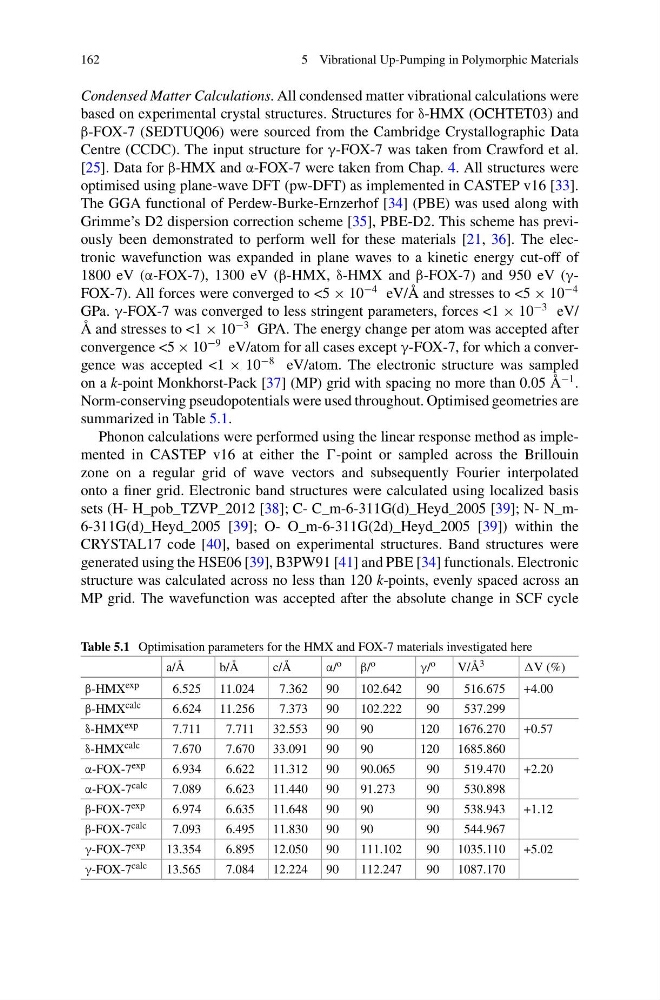
Norm-conserving pseudopotentials were used throughout. Optimised geometries are
summarized in Table 5.1.
Phonon calculations were performed using the linear response method as implemented in CASTEP v16 at either the -point or sampled across the Brillouin
zone on a regular grid of wave vectors and subsequently Fourier interpolated
onto a finer grid. Electronic band structures were calculated using localized basis
sets (H- H_pob_TZVP_2012 [38]; C- C_m-6-311G(d)_Heyd_2005 [39]; N- N_m6-311G(d)_Heyd_2005 [39]; O- O_m-6-311G(2d)_Heyd_2005 [39]) within the
CRYSTAL17 code [40], based on experimental structures. Band structures were
generated using the HSE06 [39], B3PW91 [41] and PBE [34] functionals. Electronic
structure was calculated across no less than 120 k-points, evenly spaced across an
MP grid. The wavefunction was accepted after the absolute change in SCF cycle
Table 5.1 Optimisation parameters for the HMX and FOX-7 materials investigated here
a/Å
b/Å
c/Å
α/ o β/ o
γ/ o
V/Å 3
V (%)
β-HMX exp
6.525 11.024
7.362 90
102.642
90
516.675 +4.00
β-HMX calc
6.624 11.256
7.373 90
102.222
90
537.299
δ-HMX exp
7.711
7.711 32.553 90
90
120 1676.270 +0.57
δ-HMX calc
7.670
7.670 33.091 90
90
120 1685.860
α-FOX-7 exp
6.934
6.622 11.312 90
90.065
90
519.470 +2.20
α-FOX-7 calc
7.089
6.623 11.440 90
91.273
90
530.898
β-FOX-7 exp
6.974
6.635 11.648 90
90
90
538.943 +1.12
β-FOX-7 calc
7.093
6.495 11.830 90
90
90
544.967
γ-FOX-7 exp
13.354
6.895 12.050 90
111.102
90 1035.110 +5.02
γ-FOX-7 calc
13.565
7.084 12.224 90
112.247
90 1087.170