

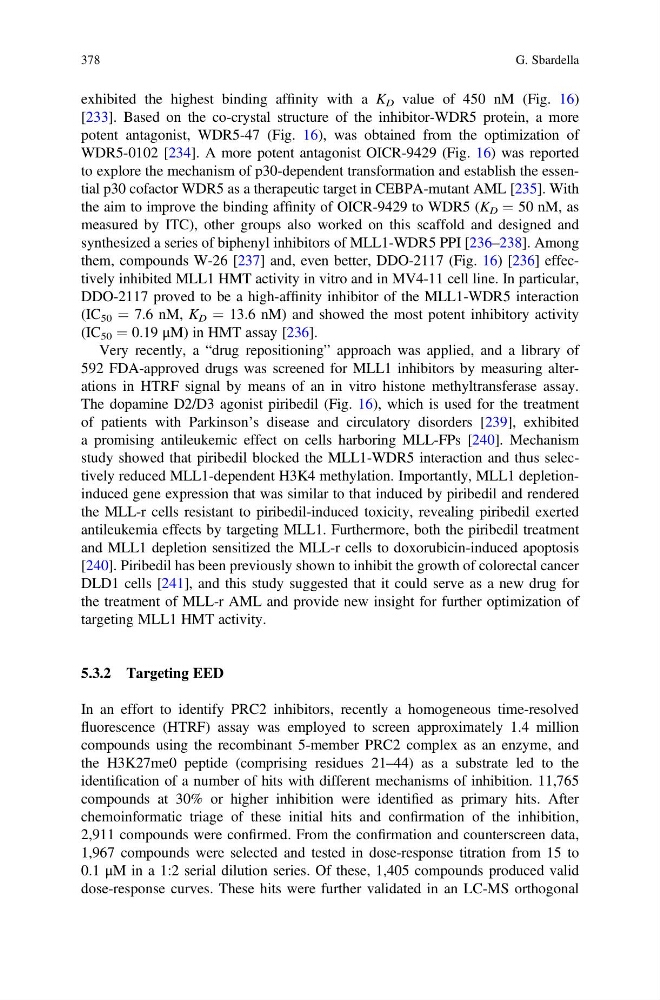
exhibited the highest binding affinity with a K D value of 450 nM (Fig. 16)
[233]. Based on the co-crystal structure of the inhibitor-WDR5 protein, a more
potent antagonist, WDR5-47 (Fig. 16), was obtained from the optimization of
WDR5-0102 [234]. A more potent antagonist OICR-9429 (Fig. 16) was reported
to explore the mechanism of p30-dependent transformation and establish the essential p30 cofactor WDR5 as a therapeutic target in CEBPA-mutant AML [235]. With
the aim to improve the binding affinity of OICR-9429 to WDR5 (K D ¼ 50 nM, as
measured by ITC), other groups also worked on this scaffold and designed and
synthesized a series of biphenyl inhibitors of MLL1-WDR5 PPI [236–238]. Among
them, compounds W-26 [237] and, even better, DDO-2117 (Fig. 16) [236] effectively inhibited MLL1 HMT activity in vitro and in MV4-11 cell line. In particular,
DDO-2117 proved to be a high-affinity inhibitor of the MLL1-WDR5 interaction
(IC 50 ¼ 7.6 nM, K D ¼ 13.6 nM) and showed the most potent inhibitory activity
(IC 50 ¼ 0.19 μM) in HMT assay [236].
Very recently, a “drug repositioning” approach was applied, and a library of
592 FDA-approved drugs was screened for MLL1 inhibitors by measuring alterations in HTRF signal by means of an in vitro histone methyltransferase assay.
The dopamine D2/D3 agonist piribedil (Fig. 16), which is used for the treatment
of patients with Parkinson’s disease and circulatory disorders [239], exhibited
a promising antileukemic effect on cells harboring MLL-FPs [240]. Mechanism
study showed that piribedil blocked the MLL1-WDR5 interaction and thus selectively reduced MLL1-dependent H3K4 methylation. Importantly, MLL1 depletioninduced gene expression that was similar to that induced by piribedil and rendered
the MLL-r cells resistant to piribedil-induced toxicity, revealing piribedil exerted
antileukemia effects by targeting MLL1. Furthermore, both the piribedil treatment
and MLL1 depletion sensitized the MLL-r cells to doxorubicin-induced apoptosis
[240]. Piribedil has been previously shown to inhibit the growth of colorectal cancer
DLD1 cells [241], and this study suggested that it could serve as a new drug for
the treatment of MLL-r AML and provide new insight for further optimization of
targeting MLL1 HMT activity.
5.3.2 Targeting EED
In an effort to identify PRC2 inhibitors, recently a homogeneous time-resolved
fluorescence (HTRF) assay was employed to screen approximately 1.4 million
compounds using the recombinant 5-member PRC2 complex as an enzyme, and
the H3K27me0 peptide (comprising residues 21–44) as a substrate led to the
identification of a number of hits with different mechanisms of inhibition. 11,765
compounds at 30% or higher inhibition were identified as primary hits. After
chemoinformatic triage of these initial hits and confirmation of the inhibition,
2,911 compounds were confirmed. From the confirmation and counterscreen data,
1,967 compounds were selected and tested in dose-response titration from 15 to
0.1 μM in a 1:2 serial dilution series. Of these, 1,405 compounds produced valid
dose-response curves. These hits were further validated in an LC-MS orthogonal
378
G. Sbardella