

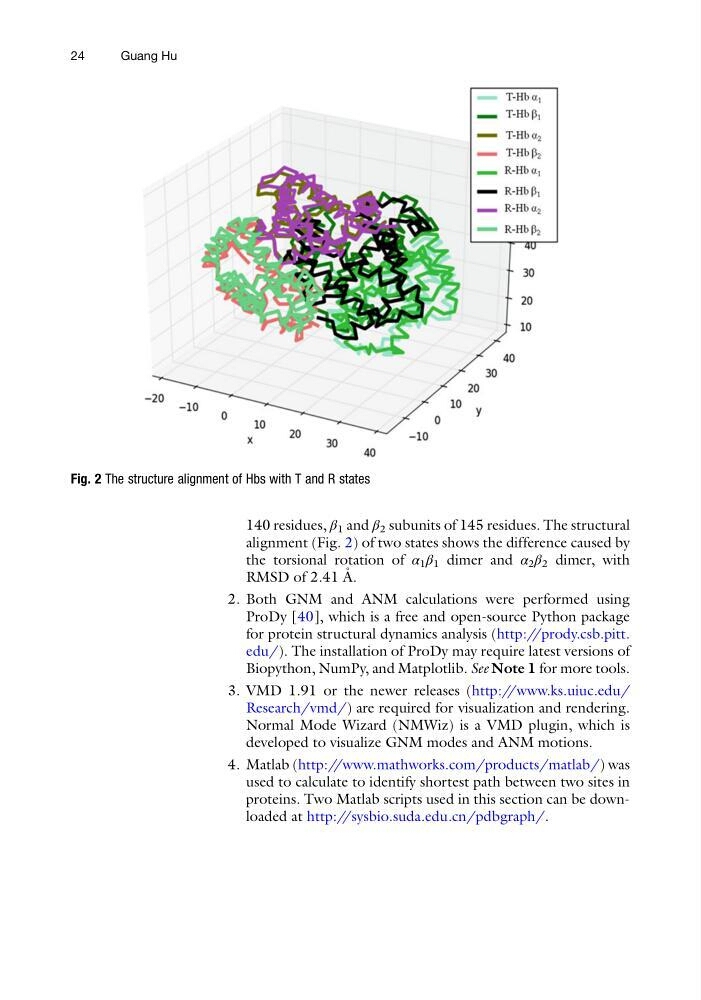
140 residues, β 1 and β 2 subunits of 145 residues. The structural
alignment (Fig. 2) of two states shows the difference caused by
the torsional rotation of α 1 β 1 dimer and α 2 β 2 dimer, with
RMSD of 2.41 A ˚ .
2. Both GNM and ANM calculations were performed using
ProDy [40], which is a free and open-source Python package
for protein structural dynamics analysis (http://prody.csb.pitt.
edu/). The installation of ProDy may require latest versions of
Biopython, NumPy, and Matplotlib. See Note 1 for more tools.
3. VMD 1.91 or the newer releases (http://www.ks.uiuc.edu/
Research/vmd/) are required for visualization and rendering.
Normal Mode Wizard (NMWiz) is a VMD plugin, which is
developed to visualize GNM modes and ANM motions.
4. Matlab (http://www.mathworks.com/products/matlab/) was
used to calculate to identify shortest path between two sites in
proteins. Two Matlab scripts used in this section can be downloaded at http://sysbio.suda.edu.cn/pdbgraph/.
Fig. 2 The structure alignment of Hbs with T and R states
24
Guang Hu