

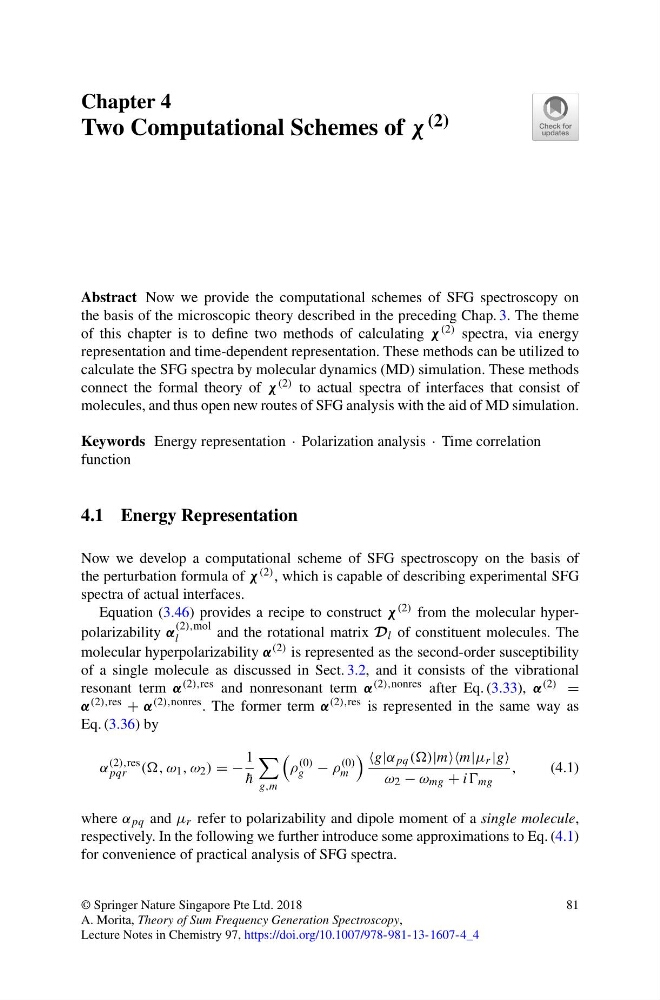
Chapter 4
Two Computational Schemes of χ (2)
Abstract Now we provide the computational schemes of SFG spectroscopy on
the basis of the microscopic theory described in the preceding Chap. 3. The theme
of this chapter is to define two methods of calculating χ (2) spectra, via energy
representation and time-dependent representation. These methods can be utilized to
calculate the SFG spectra by molecular dynamics (MD) simulation. These methods
connect the formal theory of χ (2) to actual spectra of interfaces that consist of
molecules, and thus open new routes of SFG analysis with the aid of MD simulation.
Keywords Energy representation · Polarization analysis · Time correlation
function
4.1 Energy Representation
Now we develop a computational scheme of SFG spectroscopy on the basis of
the perturbation formula of χ (2) , which is capable of describing experimental SFG
spectra of actual interfaces.
Equation (3.46) provides a recipe to construct χ (2) from the molecular hyperpolarizability α
(2),mol
l
and the rotational matrix D l of constituent molecules. The
molecular hyperpolarizability α (2) is represented as the second-order susceptibility
of a single molecule as discussed in Sect. 3.2, and it consists of the vibrational
resonant term α (2),res and nonresonant term α (2),nonres after Eq. (3.33), α (2) =
α (2),res + α (2),nonres . The former term α (2),res is represented in the same way as
Eq. (3.36) by
α
(2),res
pqr ((, ω 1 , ω 2 ) = −
1
¯
h
g,m
ρ
(0)
g − ρ
(0)
m
pq (()|m r |g
ω 2 − ω mg + ii mg
,
(4.1)
where α pq and μ r refer to polarizability and dipole moment of a single molecule,
respectively. In the following we further introduce some approximations to Eq. (4.1)
for convenience of practical analysis of SFG spectra.
© Springer Nature Singapore Pte Ltd. 2018
A. Morita, Theory of Sum Frequency Generation Spectroscopy,
Lecture Notes in Chemistry 97, https://doi.org/10.1007/978-981-13-1607-4_4
81