

258
10 Applications: Organic Interfaces
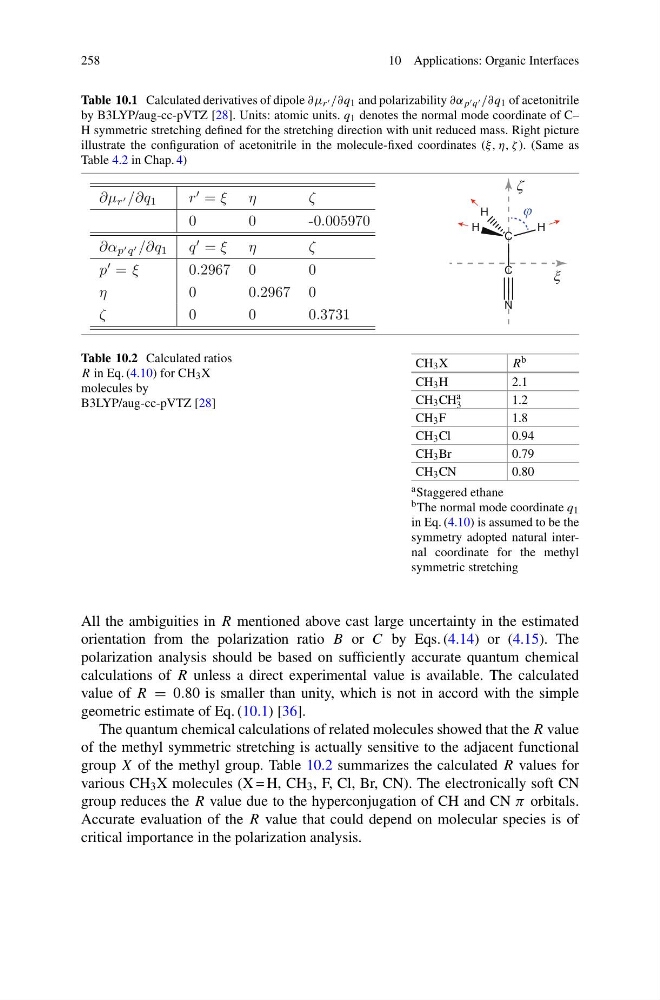
Table 10.1 Calculated derivatives of dipole ∂μ r /∂q 1 and polarizability ∂α p q /∂q 1 of acetonitrile
by B3LYP/aug-cc-pVTZ [28]. Units: atomic units. q 1 denotes the normal mode coordinate of C–
H symmetric stretching defined for the stretching direction with unit reduced mass. Right picture
illustrate the configuration of acetonitrile in the molecule-fixed coordinates (ξ, η, ζ ). (Same as
Table 4.2 in Chap. 4)
∂μ r /∂q 1
r = ξ η
ζ
0
0
-0.005970
∂α p q /∂q 1 q = ξ η
ζ
p = ξ
0.2967 0
0
η
0
0.2967 0
ζ
0
0
0.3731
C
C
H
H
H
N
Table 10.2 Calculated ratios
R in Eq. (4.10) for CH 3 X
molecules by
B3LYP/aug-cc-pVTZ [28]
CH 3 X
R b
CH 3 H
2.1
CH 3 CH
a
3
1.2
CH 3 F
1.8
CH 3 Cl
0.94
CH 3 Br
0.79
CH 3 CN
0.80
a Staggered ethane
b The normal mode coordinate q 1
in Eq. (4.10) is assumed to be the
symmetry adopted natural internal coordinate for the methyl
symmetric stretching
All the ambiguities in R mentioned above cast large uncertainty in the estimated
orientation from the polarization ratio B or C by Eqs. (4.14) or (4.15). The
polarization analysis should be based on sufficiently accurate quantum chemical
calculations of R unless a direct experimental value is available. The calculated
value of R = 0.80 is smaller than unity, which is not in accord with the simple
geometric estimate of Eq. (10.1) [36].
The quantum chemical calculations of related molecules showed that the R value
of the methyl symmetric stretching is actually sensitive to the adjacent functional
group X of the methyl group. Table 10.2 summarizes the calculated R values for
various CH 3 X molecules (X = H, CH 3 , F, Cl, Br, CN). The electronically soft CN
group reduces the R value due to the hyperconjugation of CH and CN π orbitals.
Accurate evaluation of the R value that could depend on molecular species is of
critical importance in the polarization analysis.