

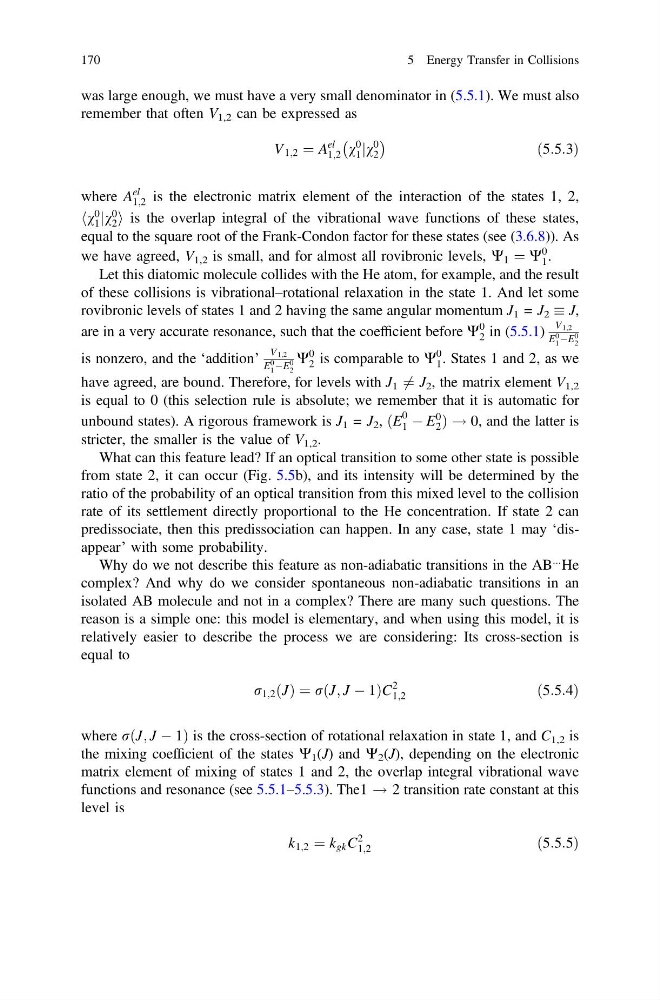
was large enough, we must have a very small denominator in (5.5.1). We must also
remember that often V 1,2 can be expressed as
V 1;2 ¼ A
el
1;2 v
0
1 jv
0
2
À
Á
ð5:5:3Þ
where A
el
1;2 is the electronic matrix element of the interaction of the states 1, 2,
hv
0
1 jv
0
2 i is the overlap integral of the vibrational wave functions of these states,
equal to the square root of the Frank-Condon factor for these states (see (3.6.8)). As
we have agreed, V 1,2 is small, and for almost all rovibronic levels, W 1 ¼ W
0
1 .
Let this diatomic molecule collides with the He atom, for example, and the result
of these collisions is vibrational–rotational relaxation in the state 1. And let some
rovibronic levels of states 1 and 2 having the same angular momentum J 1 = J 2 J,
are in a very accurate resonance, such that the coefficient before W
0
2 in (5.5.1)
V 1;2
E
0
1 ÀE
0
2
is nonzero, and the ‘addition’
V 1;2
E
0
1 ÀE
0
2
W
0
2 is comparable to W
0
1 . States 1 and 2, as we
have agreed, are bound. Therefore, for levels with J 1 6 ¼ J 2 , the matrix element V 1,2
is equal to 0 (this selection rule is absolute; we remember that it is automatic for
unbound states). A rigorous framework is J 1 = J 2 , ðE
0
1 À E
0
2 Þ ! 0, and the latter is
stricter, the smaller is the value of V 1,2 .
What can this feature lead? If an optical transition to some other state is possible
from state 2, it can occur (Fig. 5.5b), and its intensity will be determined by the
ratio of the probability of an optical transition from this mixed level to the collision
rate of its settlement directly proportional to the He concentration. If state 2 can
predissociate, then this predissociation can happen. In any case, state 1 may ‘disappear’ with some probability.
Why do we not describe this feature as non-adiabatic transitions in the AB
... He
complex? And why do we consider spontaneous non-adiabatic transitions in an
isolated AB molecule and not in a complex? There are many such questions. The
reason is a simple one: this model is elementary, and when using this model, it is
relatively easier to describe the process we are considering: Its cross-section is
equal to
r 1;2 ðJÞ ¼ rðJ; J À 1ÞC
2
1;2
ð5:5:4Þ
where rðJ; J À 1Þ is the cross-section of rotational relaxation in state 1, and C 1,2 is
the mixing coefficient of the states W 1 (J) and W 2 (J), depending on the electronic
matrix element of mixing of states 1 and 2, the overlap integral vibrational wave
functions and resonance (see 5.5.1–5.5.3). The1 ! 2 transition rate constant at this
level is
k 1;2 ¼ k gk C
2
1;2
ð5:5:5Þ
170
5 Energy Transfer in Collisions