

et al. 2016; Klocko et al. 2019), as observed for
many other organisms (Lai and Pugh 2017).
Additional accessible regions within intergenic
sequences can also be identified, presumably
corresponding to important distal regulatory
sequences that control transcription of a nearby
promoter. On the other hand, well-positioned
nucleosomes can be observed adjacent to the
NFR extending into the gene coding sequence
(see Sect. II.B).
Several mechanisms operate to create or
maintain open chromatin within gene promoters. In other eukaryotes, promoter regions
adjacent to the transcriptional start site are
enriched for sequences that are energetically
unfavorable for nucleosome binding, such as
poly(dA:dT) tracts (Iyer and Struhl 1995; Segal
and Widom 2009; Krietenstein et al. 2016). It is
not known if this also occurs in N. crassa, but
given the similarity in nucleosome occupancy
pattern surrounding transcriptional start sites
(i.e., an NFR is often observed just upstream of
the transcriptional start site), it is reasonable
that the NFRs of N. crassa may be enriched for
similar sequences. Other mechanisms that promote chromatin accessibility involve regulatory
proteins or protein complexes. ATP-dependent
chromatin remodeling enzymes can function
to enhance chromatin accessibility, for example. The major class of ATP-dependent remodeling enzymes is the SWI/SNF (switching
defective/sucrose nonfermenting) superfamily,
named after the phenotypes of yeast mutants
that led to the discovery of the founding family
member (Stern et al. 1984; Neigeborn and Carlson 1984). SWI/SNF family remodeling
enzymes harness the energy from ATP hydrolysis to alter local chromatin structure by
changing the absolute position or composition
of the histone octamer on DNA through a variety of mechanisms including sliding nucleosomes along the DNA, ejecting nucleosomes
from the DNA, ejecting histone dimers from
the histone octamer, and, in the case of the
multi-subunit SWR1 complex, depositing the
histone variant H2A.Z in place of the canonical
H2A (Clapier and Cairns 2009; Clapier et al.
2017). These activities can increase or decrease
chromatin accessibility. A number of N. crassa
remodelers have been implicated in gene regulation, primarily linked to the regulation of
circadian clock genes. For example, the SNF2
family enzyme CLOCKSWITCH (NCU09106)
and the multi-subunit SWI/SNF complex were
shown to remodel chromatin in the promoter
and upstream regulatory sequences controlling
frequency (frq; NCU02265), a key component of
circadian oscillator (Belden et al. 2007; Wang
et al. 2014). Another chromatin remodeler, the
Clock-ATPase (CATP; NCU06484), is necessary
to displace histones from the C-box, a critical
cis-regulatory element required for activation
of frq transcription. Disruption of the catp
gene caused increased histone H3 density at
the C-box, reduced frq transcription, and an
aberrant circadian rhythm (Cha et al. 2013).
Interestingly, the catp gene was also identified
as the defective in methylation mutant dim-1
locus (Sect. IIIA) (Klocko et al. 2019), suggesting this chromatin remodeler acts across the
genome at both euchromatin and heterochromatin.
Another major mechanism for creating
“open” chromatin is the binding of transcription factors. Nucleosome occupancy and transcription factor binding are typically mutually
exclusive, such that transcription factors can
compete with nucleosomes for access to the
underlying DNA at promoters. In N. crassa,
the DNA-binding photoreceptor White Collar
Complex (WCC), comprised of WC1
[NCU02356] and WC2 [NCU00902], is required
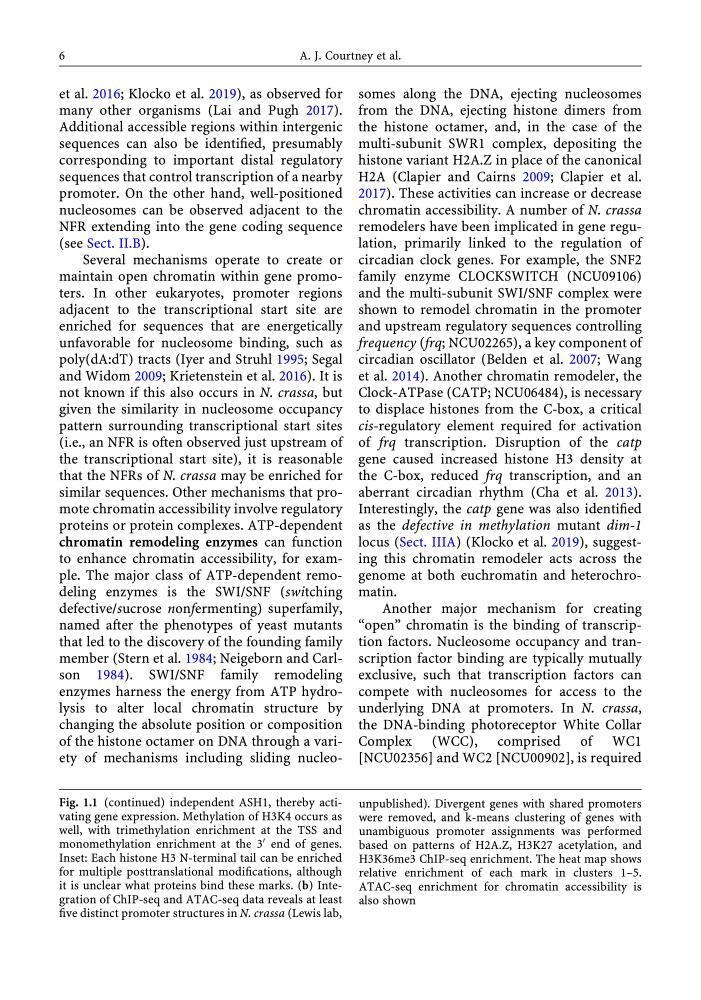
Fig. 1.1 (continued) independent ASH1, thereby activating gene expression. Methylation of H3K4 occurs as
well, with trimethylation enrichment at the TSS and
monomethylation enrichment at the 3
0 end of genes.
Inset: Each histone H3 N-terminal tail can be enriched
for multiple posttranslational modifications, although
it is unclear what proteins bind these marks. (b) Integration of ChIP-seq and ATAC-seq data reveals at least
five distinct promoter structures in N. crassa (Lewis lab,
unpublished). Divergent genes with shared promoters
were removed, and k-means clustering of genes with
unambiguous promoter assignments was performed
based on patterns of H2A.Z, H3K27 acetylation, and
H3K36me3 ChIP-seq enrichment. The heat map shows
relative enrichment of each mark in clusters 1–5.
ATAC-seq enrichment for chromatin accessibility is
also shown
6
A. J. Courtney et al.