

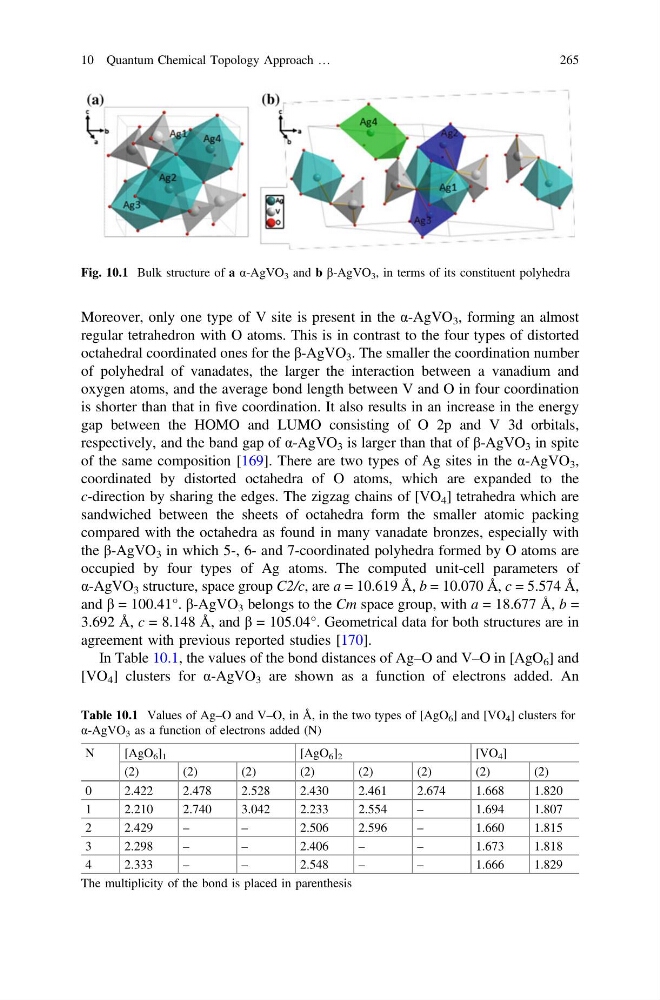
Moreover, only one type of V site is present in the α-AgVO 3 , forming an almost
regular tetrahedron with O atoms. This is in contrast to the four types of distorted
octahedral coordinated ones for the β-AgVO 3 . The smaller the coordination number
of polyhedral of vanadates, the larger the interaction between a vanadium and
oxygen atoms, and the average bond length between V and O in four coordination
is shorter than that in five coordination. It also results in an increase in the energy
gap between the HOMO and LUMO consisting of O 2p and V 3d orbitals,
respectively, and the band gap of α-AgVO 3 is larger than that of β-AgVO 3 in spite
of the same composition [169]. There are two types of Ag sites in the α-AgVO 3 ,
coordinated by distorted octahedra of O atoms, which are expanded to the
c-direction by sharing the edges. The zigzag chains of [VO 4 ] tetrahedra which are
sandwiched between the sheets of octahedra form the smaller atomic packing
compared with the octahedra as found in many vanadate bronzes, especially with
the β-AgVO 3 in which 5-, 6- and 7-coordinated polyhedra formed by O atoms are
occupied by four types of Ag atoms. The computed unit-cell parameters of
α-AgVO 3 structure, space group C2/c, are a = 10.619 Å, b = 10.070 Å, c = 5.574 Å,
and β = 100.41°. β-AgVO 3 belongs to the Cm space group, with a = 18.677 Å, b =
3.692 Å, c = 8.148 Å, and β = 105.04°. Geometrical data for both structures are in
agreement with previous reported studies [170].
In Table 10.1, the values of the bond distances of Ag–O and V–O in [AgO 6 ] and
[VO 4 ] clusters for α-AgVO 3 are shown as a function of electrons added. An
Fig. 10.1 Bulk structure of a α-AgVO 3 and b β-AgVO 3 , in terms of its constituent polyhedra
Table 10.1 Values of Ag–O and V–O, in Å, in the two types of [AgO 6 ] and [VO 4 ] clusters for
α-AgVO 3 as a function of electrons added (N)
N
[AgO 6 ] 1
[AgO 6 ] 2
[VO 4 ]
(2)
(2)
(2)
(2)
(2)
(2)
(2)
(2)
0
2.422
2.478
2.528
2.430
2.461
2.674
1.668
1.820
1
2.210
2.740
3.042
2.233
2.554
–
1.694
1.807
2
2.429
–
–
2.506
2.596
–
1.660
1.815
3
2.298
–
–
2.406
–
–
1.673
1.818
4
2.333
–
–
2.548
–
–
1.666
1.829
The multiplicity of the bond is placed in parenthesis
10 Quantum Chemical Topology Approach …
265