

464
N. Ya. Golovenko et al.
Where D is the diffusivity of the dissolved drug, ρ is the density of the dissolved
drug, C S is the drug solubility, and r is the initial radius of the drug particle.
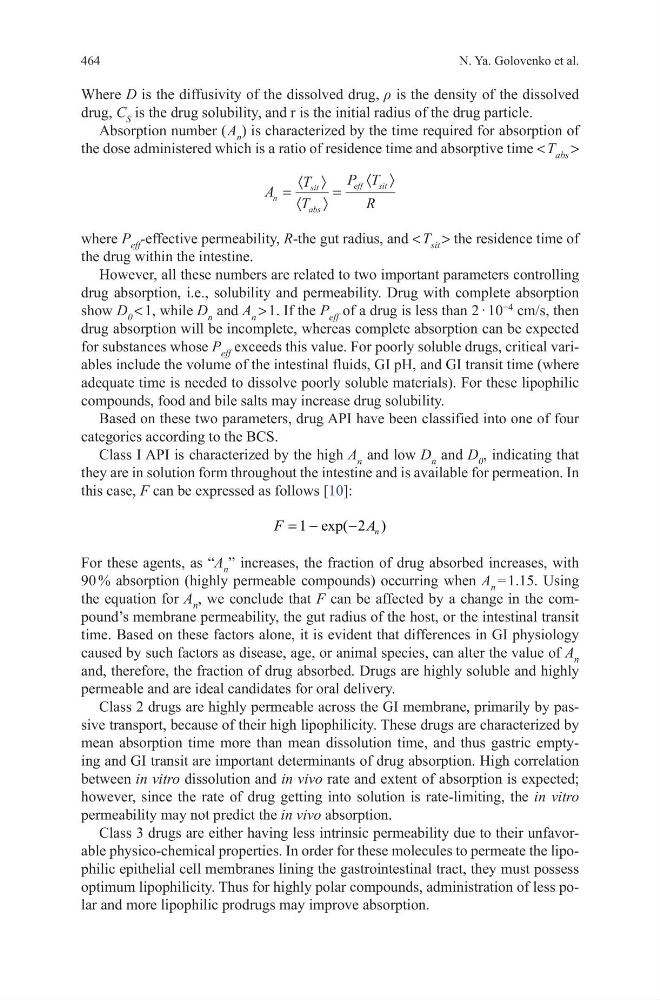
Absorption number ( A n ) is characterized by the time required for absorption of
the dose administered which is a ratio of residence time and absorptive time < T abs >
where P eff -effective permeability, R-the gut radius, and < T sit > the residence time of
the drug within the intestine.
However, all these numbers are related to two important parameters controlling
drug absorption, i.e., solubility and permeability. Drug with complete absorption
show D 0 < 1, while D n and A n > 1. If the P eff of a drug is less than 2 · 10
−4
cm/s, then
drug absorption will be incomplete, whereas complete absorption can be expected
for substances whose P eff exceeds this value. For poorly soluble drugs, critical variables include the volume of the intestinal fluids, GI pH, and GI transit time (where
adequate time is needed to dissolve poorly soluble materials). For these lipophilic
compounds, food and bile salts may increase drug solubility.
Based on these two parameters, drug API have been classified into one of four
categories according to the BCS.
Class I API is characterized by the high A n and low D n and D 0 , indicating that
they are in solution form throughout the intestine and is available for permeation. In
this case, F can be expressed as follows [10]:
For these agents, as “A n ” increases, the fraction of drug absorbed increases, with
90 % absorption (highly permeable compounds) occurring when A n = 1.15. Using
the equation for A n , we conclude that F can be affected by a change in the compound’s membrane permeability, the gut radius of the host, or the intestinal transit
time. Based on these factors alone, it is evident that differences in GI physiology
caused by such factors as disease, age, or animal species, can alter the value of A n
and, therefore, the fraction of drug absorbed. Drugs are highly soluble and highly
permeable and are ideal candidates for oral delivery.
Class 2 drugs are highly permeable across the GI membrane, primarily by passive transport, because of their high lipophilicity. These drugs are characterized by
mean absorption time more than mean dissolution time, and thus gastric emptying and GI transit are important determinants of drug absorption. High correlation
between in vitro dissolution and in vivo rate and extent of absorption is expected;
however, since the rate of drug getting into solution is rate-limiting, the in vitro
permeability may not predict the in vivo absorption.
Class 3 drugs are either having less intrinsic permeability due to their unfavorable physico-chemical properties. In order for these molecules to permeate the lipophilic epithelial cell membranes lining the gastrointestinal tract, they must possess
optimum lipophilicity. Thus for highly polar compounds, administration of less polar and more lipophilic prodrugs may improve absorption.
A
T
T
P T
R
n
sit
abs
eff
s it
=
〈 〉
〈 〉
=
〈 〉
1 exp( 2 )
n
F
A
= -
-