

3.1 Structured Bodies
91
further higher levels of theory have been built. The outcomes of these higher level
of theory calculations will instead be taken as electronic structure values to fit.
3.2 Higher Level Ab initio Methods
3.2.1 Beyond the Hartree–Fock Method
As already mentioned, merits and demerits of HF calculations have been repeatedly
analyzed in the literature to the end of devising corrective actions. In fact, while
the HF total energy represents more than 90% of the nonrelativistic one the differences in energy coming into play in chemical reactions may be of the order of
100 or 1000 times smaller. As a matter of fact when a chemical process implies the
breaking/forming or even the simple exploration of potential energy regions ranging
from long to short distances (with the consequent strong perturbation of the electronic distribution) one has to incorporate in the ab initio calculation the electronic
correlation.
Let us start with a list of peculiarities of the HF method:
• the application of the variational principle and of the orthogonality one meets the
basic accuracy criteria only for the ground state
• the SCF nature of the HF method guarantees that the electrostatic potential
described by the solution wavefunctions is the same as that of the operator
• the wavefunctions not appearing in the final composition of the operator are virtual
• the energy of the HF determinant differs from that of the orbitals; their sum is the
energy corrected to the next order
• Open and closed shell HF solutions are different and require two different ways
(unrestricted and restricted) of dealing with the total spin.
A first point to make is that the original Fock equations were designed for atomic
systems (that is for systems of spherical symmetry). In 1951 Hall and Roothaan [21]
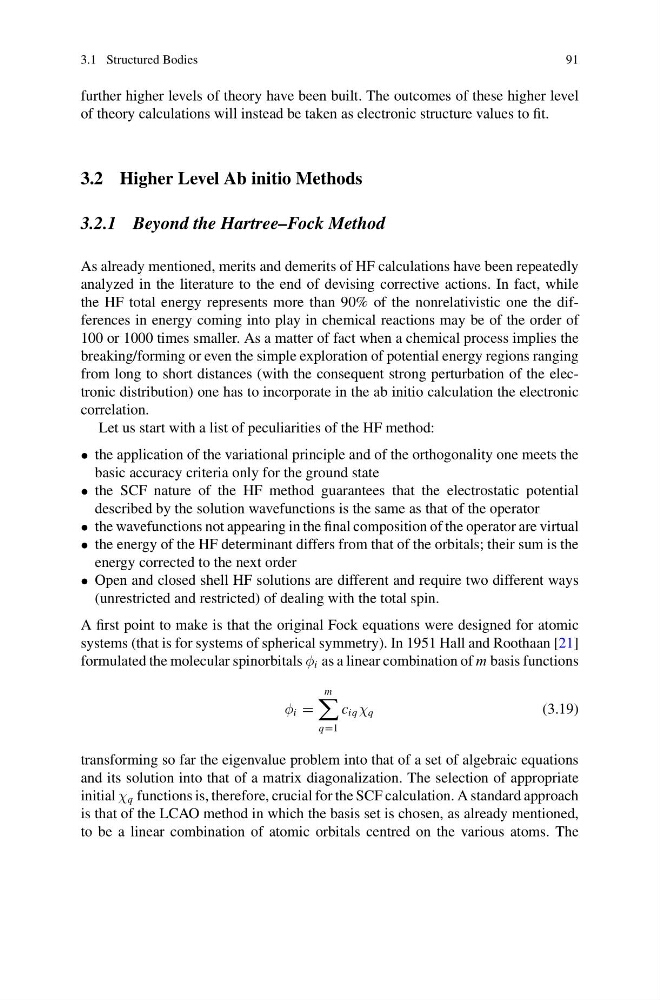
formulated the molecular spinorbitals φ i as a linear combination of m basis functions
φ i =
m
q=1
c iq χ q
(3.19)
transforming so far the eigenvalue problem into that of a set of algebraic equations
and its solution into that of a matrix diagonalization. The selection of appropriate
initial χ q functions is, therefore, crucial for the SCF calculation. A standard approach
is that of the LCAO method in which the basis set is chosen, as already mentioned,
to be a linear combination of atomic orbitals centred on the various atoms. The