

Theor Chem Acc (2015) 134:86
1 3
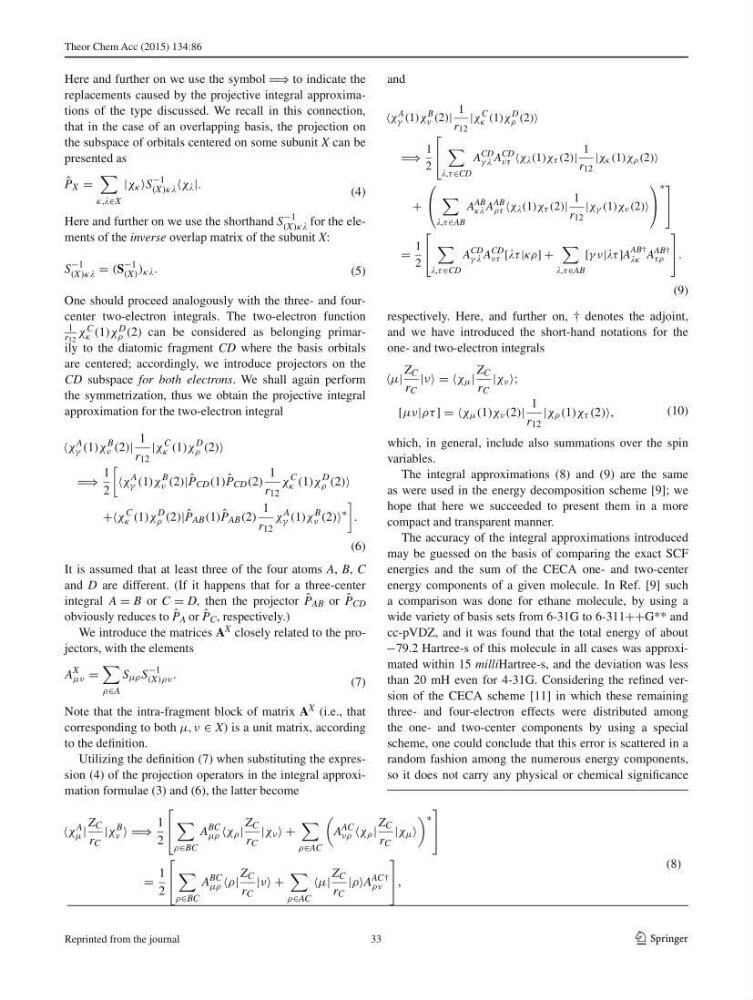
Here and further on we use the symbol =⇒ to indicate the
replacements caused by the projective integral approximations of the type discussed. We recall in this connection,
that in the case of an overlapping basis, the projection on
the subspace of orbitals centered on some subunit X can be
presented as
Here and further on we use the shorthand S
−1
(X)κ for the elements of the inverse overlap matrix of the subunit X :
One should proceed analogously with the three- and fourcenter two-electron integrals. The two-electron function
1
r 12
χ C
κ (1)χ D
ρ (2) can be considered as belonging primarily to the diatomic fragment CD where the basis orbitals
are centered; accordingly, we introduce projectors on the
CD subspace for both electrons . We shall again perform
the symmetrization, thus we obtain the projective integral
approximation for the two-electron integral
It is assumed that at least three of the four atoms A , B , C
and D are different. (If it happens that for a three-center
integral A = B or C = D , then the projector ˆ
P AB or ˆ
P CD
obviously reduces to ˆ
P A or ˆ
P C , respectively.)
We introduce the matrices A X closely related to the projectors, with the elements
Note that the intra-fragment block of matrix A X (i.e., that
corresponding to both μ, ν ∈ X ) is a unit matrix, according
to the defi nition.
Utilizing the defi nition ( 7 ) when substituting the expression ( 4 ) of the projection operators in the integral approximation formulae ( 3 ) and ( 6 ), the latter become
(4)
ˆ
P X =
κ,∈X
|χ κ S
−1
(X)κ χ |.
(5)
S
−1
(X)κ = (S
−1
(X) ) κ .
(6)
χ
A
γ (1)χ
B
ν (2)|
1
r 12
|χ
C
κ (1)χ
D
ρ (2)
=⇒
1
2
χ
A
γ (1)χ
B
ν (2)| ˆ
P CD (1) ˆ
P CD (2)
1
r 12
χ
C
κ (1)χ
D
ρ (2)
++χ
C
κ (1)χ
D
ρ (2)| ˆ
P AB (1) ˆ
P AB (2)
1
r 12
χ
A
γ (1)χ
B
ν (2)
∗
.
(7)
A
X
μν =
ρ∈A
S μρ S
−1
(X)ρν .
(8)
χ
A
μ |
Z C
r C
|χ
B
ν =⇒
1
2
⎡
⎣
ρ∈BC
A
BC
μρ χ ρ |
Z C
r C
|χ ν +
ρ∈AC
A
AC
νρ χ ρ |
Z C
r C
|χ μ
∗
⎤
⎦
=
1
2
⎡
⎣
ρ∈BC
A
BC
μρ ρ|
Z C
r C
|ν +
ρ∈AC
μ|
Z C
r C
|ρA
AC†
ρν
⎤
⎦ ,
and
respectively. Here, and further on, † denotes the adjoint,
and we have introduced the short-hand notations for the
one- and two-electron integrals
which, in general, include also summations over the spin
variables.
The integral approximations ( 8 ) and ( 9 ) are the same
as were used in the energy decomposition scheme [ 9 ]; we
hope that here we succeeded to present them in a more
compact and transparent manner.
The accuracy of the integral approximations introduced
may be guessed on the basis of comparing the exact SCF
energies and the sum of the CECA one- and two-center
energy components of a given molecule. In Ref. [ 9 ] such
a comparison was done for ethane molecule, by using a
wide variety of basis sets from 6-31G to 6-311++G** and
cc-pVDZ, and it was found that the total energy of about
−79.2 Hartree-s of this molecule in all cases was approximated within 15 milli Hartree-s, and the deviation was less
than 20 mH even for 4-31G. Considering the refi ned version of the CECA scheme [ 11 ] in which these remaining
three- and four-electron effects were distributed among
the one- and two-center components by using a special
scheme, one could conclude that this error is scattered in a
random fashion among the numerous energy components,
so it does not carry any physical or chemical signifi cance
(9)
χ
A
γ (1)χ
B
ν (2)|
1
r 12
|χ
C
κ (1)χ
D
ρ (2)
=⇒
1
2
⎡
⎣
,τ ∈CD
A
CD
γ A
CD
ντ χ (1)χ τ (2)|
1
r 12
|χ κ (1)χ ρ (2)
+
⎛
⎝
,τ ∈AB
A
AB
κ A
AB
ρτ χ (1)χ τ (2)|
1
r 12
|χ γ (1)χ ν (2)
⎞
⎠
∗ ⎤
⎦
=
1
2
⎡
⎣
,τ ∈CD
A
CD
γ A
CD
ντ [τ |κρ] +
,τ ∈AB
[γ ν|τ ]A
AB†
κ A
AB†
τρ
⎤
⎦ .
(10)
μ|
Z C
r C
|ν = =χ μ |
Z C
r C
|χ ν ;
[μν|ρτ ] = =χ μ (1)χ ν (2)|
1
r 12
|χ ρ (1)χ τ (2),
33
Reprinted from the journal