

Theor Chem Acc (2015) 134:116
1 3
aqueous-ethanolic solution and characterized by X-ray
structure analysis and spectral methods (infrared (IR),
Raman, UV–Vis,
51 V NMR) [ 16 , 17 ] were included in our
computational study:
While the tetranuclear anion [ 17 ] always contains tartrato groups of the same enantiomeric form, formation
of the dinuclear complexes is stereospecifi c [ 16 ]. Continuous effort to understand the stereospecifi c aspects of
vanadium(V) tartrato complexes leads us to take a closer
look at their spectral properties. Hence, this work is aimed
to complement the aforementioned studies and provide
deeper understanding of the experimental observations.
At present, the density functional theory (DFT) prevails
in calculations of the structural and spectral parameters of
transition metal complexes [ 18 ]. For this reason, and thanks
to our previous positive experience with simulation and
interpretation of the UV–Vis, IR, electronic and vibrational
circular dichroism (ECD and VCD) spectra of chiral vanadium complexes [ 19 ], we decided to use the DFT methods
as the main tool in the present work as well. We shall present the calculated molecular structures, vibrational and
electronic spectra, as well as the
51 V chemical shifts of
the chosen anions of the vanadium(V) tartrato complexes.
Where applicable, results are confronted with the available
experimental data, with the aim to assess the reliability of
the individual methods for future calculations of a similar
kind.
2 Experimental details
Absorption UV–Vis and ECD spectra were recorded on a
JASCO J-815 CD spectrometer in CH 3 CN and H 2 O solutions with a 1-cm cell.
Experimentally observed IR and Raman fundamentals
were extracted from the previously published spectra, and
compounds were prepared based on established synthetic
procedure [ 16 , 17 ].
3 Computational details
The quantum chemistry calculations were carried out using
Gaussian 09 software package [ 20 ]. For visualization of
the molecular structures and assignments of the vibrational
modes, the Molden program [ 21 ] was used.
Our largest complex anion includes 36 atoms and altogether 308 electrons, and the basis sets have been chosen
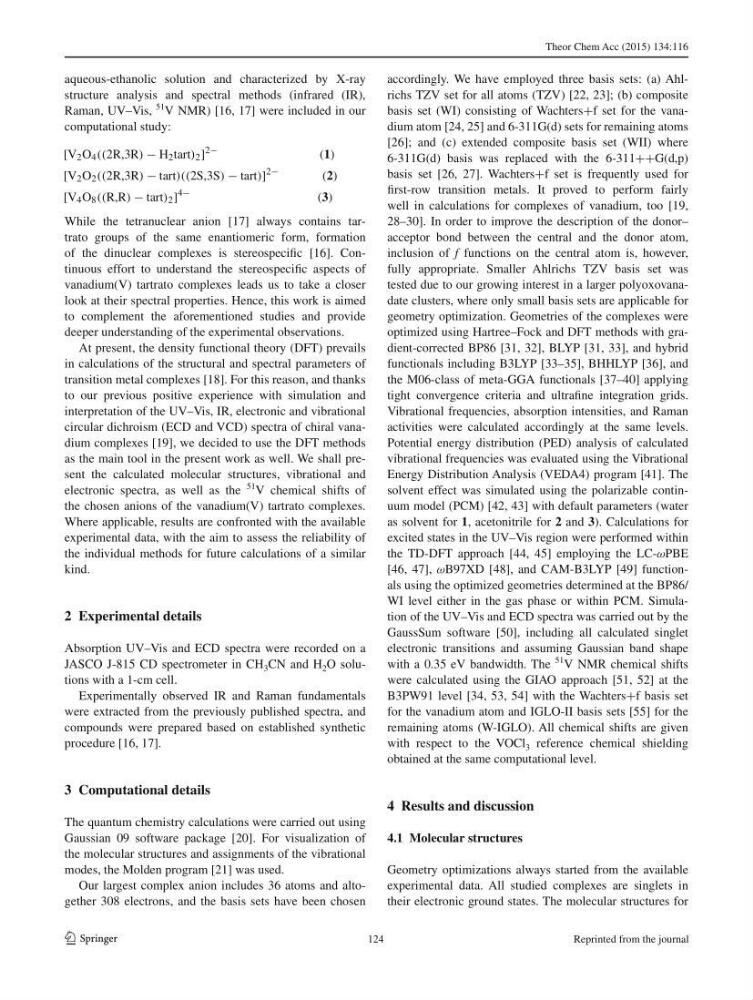
[V 2 O 4 ((2R,3R) − H 2 tart) 2 ]
2−
(1)
[V 2 O 2 ((2R,3R) − tart)((2S,3S) − tart)]
2−
(2)
[V 4 O 8 ((R,R) − tart) 2 ]
4−
(3)
accordingly. We have employed three basis sets: (a) Ahlrichs TZV set for all atoms (TZV) [ 22 , 23 ]; (b) composite
basis set (WI) consisting of Wachters+f set for the vanadium atom [ 24 , 25 ] and 6-311G(d) sets for remaining atoms
[ 26 ]; and (c) extended composite basis set (WII) where
6-311G(d) basis was replaced with the 6-311++G(d,p)
basis set [ 26 , 27 ]. Wachters+f set is frequently used for
fi rst-row transition metals. It proved to perform fairly
well in calculations for complexes of vanadium, too [ 19 ,
28 – 30 ]. In order to improve the description of the donor–
acceptor bond between the central and the donor atom,
inclusion of f functions on the central atom is, however,
fully appropriate. Smaller Ahlrichs TZV basis set was
tested due to our growing interest in a larger polyoxovanadate clusters, where only small basis sets are applicable for
geometry optimization. Geometries of the complexes were
optimized using Hartree–Fock and DFT methods with gradient-corrected BP86 [ 31 , 32 ], BLYP [ 31 , 33 ], and hybrid
functionals including B3LYP [ 33 – 35 ], BHHLYP [ 36 ], and
the M06-class of meta-GGA functionals [ 37 – 40 ] applying
tight convergence criteria and ultrafi ne integration grids.
Vibrational frequencies, absorption intensities, and Raman
activities were calculated accordingly at the same levels.
Potential energy distribution (PED) analysis of calculated
vibrational frequencies was evaluated using the Vibrational
Energy Distribution Analysis (VEDA4) program [ 41 ]. The
solvent effect was simulated using the polarizable continuum model (PCM) [ 42 , 43 ] with default parameters (water
as solvent for 1 , acetonitrile for 2 and 3 ). Calculations for
excited states in the UV–Vis region were performed within
the TD-DFT approach [ 44 , 45 ] employing the LC- ω PBE
[ 46 , 47 ], ω B97XD [ 48 ], and CAM-B3LYP [ 49 ] functionals using the optimized geometries determined at the BP86/
WI level either in the gas phase or within PCM. Simulation of the UV–Vis and ECD spectra was carried out by the
GaussSum software [ 50 ], including all calculated singlet
electronic transitions and assuming Gaussian band shape
with a 0.35 eV bandwidth. The
51 V NMR chemical shifts
were calculated using the GIAO approach [ 51 , 52 ] at the
B3PW91 level [ 34 , 53 , 54 ] with the Wachters+f basis set
for the vanadium atom and IGLO-II basis sets [ 55 ] for the
remaining atoms (W-IGLO). All chemical shifts are given
with respect to the VOCl 3 reference chemical shielding
obtained at the same computational level.
4 Results and discussion
4.1 Molecular structures
Geometry optimizations always started from the available
experimental data. All studied complexes are singlets in
their electronic ground states. The molecular structures for
124
Reprinted from the journal