

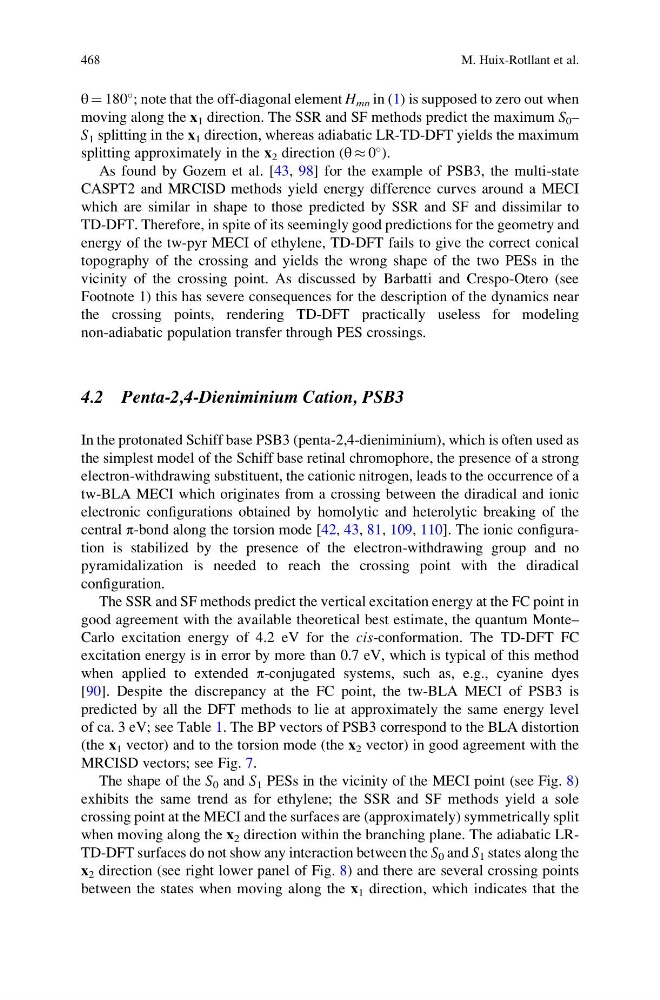
θ ¼ 180
; note that the off-diagonal element H mn in (1) is supposed to zero out when
moving along the x 1 direction. The SSR and SF methods predict the maximum S 0 –
S 1 splitting in the x 1 direction, whereas adiabatic LR-TD-DFT yields the maximum
splitting approximately in the x 2 direction (θ % 0
).
As found by Gozem et al. [43, 98] for the example of PSB3, the multi-state
CASPT2 and MRCISD methods yield energy difference curves around a MECI
which are similar in shape to those predicted by SSR and SF and dissimilar to
TD-DFT. Therefore, in spite of its seemingly good predictions for the geometry and
energy of the tw-pyr MECI of ethylene, TD-DFT fails to give the correct conical
topography of the crossing and yields the wrong shape of the two PESs in the
vicinity of the crossing point. As discussed by Barbatti and Crespo-Otero (see
Footnote 1) this has severe consequences for the description of the dynamics near
the crossing points, rendering TD-DFT practically useless for modeling
non-adiabatic population transfer through PES crossings.
4.2 Penta-2,4-Dieniminium Cation, PSB3
In the protonated Schiff base PSB3 (penta-2,4-dieniminium), which is often used as
the simplest model of the Schiff base retinal chromophore, the presence of a strong
electron-withdrawing substituent, the cationic nitrogen, leads to the occurrence of a
tw-BLA MECI which originates from a crossing between the diradical and ionic
electronic configurations obtained by homolytic and heterolytic breaking of the
central π-bond along the torsion mode [42, 43, 81, 109, 110]. The ionic configuration is stabilized by the presence of the electron-withdrawing group and no
pyramidalization is needed to reach the crossing point with the diradical
configuration.
The SSR and SF methods predict the vertical excitation energy at the FC point in
good agreement with the available theoretical best estimate, the quantum Monte–
Carlo excitation energy of 4.2 eV for the cis-conformation. The TD-DFT FC
excitation energy is in error by more than 0.7 eV, which is typical of this method
when applied to extended π-conjugated systems, such as, e.g., cyanine dyes
[90]. Despite the discrepancy at the FC point, the tw-BLA MECI of PSB3 is
predicted by all the DFT methods to lie at approximately the same energy level
of ca. 3 eV; see Table 1. The BP vectors of PSB3 correspond to the BLA distortion
(the x 1 vector) and to the torsion mode (the x 2 vector) in good agreement with the
MRCISD vectors; see Fig. 7.
The shape of the S 0 and S 1 PESs in the vicinity of the MECI point (see Fig. 8)
exhibits the same trend as for ethylene; the SSR and SF methods yield a sole
crossing point at the MECI and the surfaces are (approximately) symmetrically split
when moving along the x 2 direction within the branching plane. The adiabatic LRTD-DFT surfaces do not show any interaction between the S 0 and S 1 states along the
x 2 direction (see right lower panel of Fig. 8) and there are several crossing points
between the states when moving along the x 1 direction, which indicates that the
468
M. Huix-Rotllant et al.