

with catalyst 9/MAO are about half those of the corresponding rmmr pentad
concentrations determined for polymers produced with catalysts of complexes
(1 and 6)/MAO. Additionally, rmmr pentads in syndiotactic polymers produced
with 9/MAO show an even more moderate temperature dependency than the rmmr
temperature dependency observed for polymers of (1 and 6)/MAO.
The improved enantioselectivity, reflected in lower percentage of rmmr pentads,
for catalyst system 9/MAO could be explained by the beneficial presence/action of
the bulky tert-butyl substituents placed at positions 3 and 6 of the fluorenyl section
of the ligand. These direct the polymer chain to the most preferred “upward”
conformation away from fluorenyl part of the ligand and toward available quadrants
left or right of the cyclopentadienyl group (see Sect. 3.2) and, at the same time,
provide a more effective guidance for the preferred head-down propylene
coordination mode (anti position with respect to the polymer chain via creation
of a tighter “chiral pocket”). The explanation for the lower site epimerization
related rrmr pentad concentration in polymers produced with 9/MAO versus
1/MAO, at the same polymerization temperature and monomer concentration is,
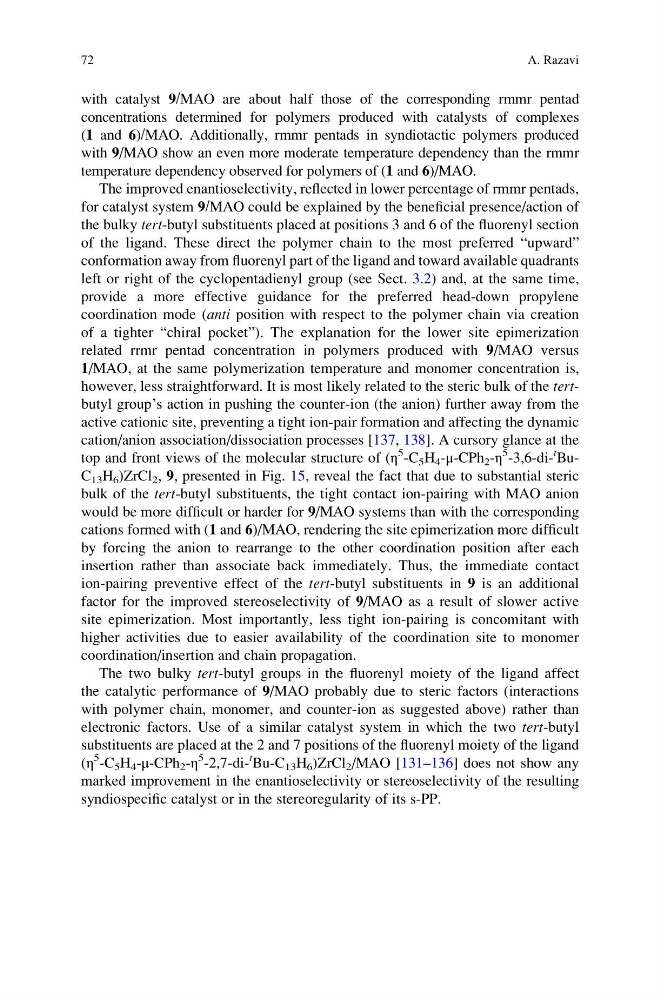
however, less straightforward. It is most likely related to the steric bulk of the tertbutyl group’s action in pushing the counter-ion (the anion) further away from the
active cationic site, preventing a tight ion-pair formation and affecting the dynamic
cation/anion association/dissociation processes [137, 138]. A cursory glance at the
top and front views of the molecular structure of (η
5 -C 5 H 4 -μ-CPh 2 -η
5 -3,6-dit
BuC 13 H 6 )ZrCl 2 , 9, presented in Fig. 15, reveal the fact that due to substantial steric
bulk of the tert-butyl substituents, the tight contact ion-pairing with MAO anion
would be more difficult or harder for 9/MAO systems than with the corresponding
cations formed with (1 and 6)/MAO, rendering the site epimerization more difficult
by forcing the anion to rearrange to the other coordination position after each
insertion rather than associate back immediately. Thus, the immediate contact
ion-pairing preventive effect of the tert-butyl substituents in 9 is an additional
factor for the improved stereoselectivity of 9/MAO as a result of slower active
site epimerization. Most importantly, less tight ion-pairing is concomitant with
higher activities due to easier availability of the coordination site to monomer
coordination/insertion and chain propagation.
The two bulky tert-butyl groups in the fluorenyl moiety of the ligand affect
the catalytic performance of 9/MAO probably due to steric factors (interactions
with polymer chain, monomer, and counter-ion as suggested above) rather than
electronic factors. Use of a similar catalyst system in which the two tert-butyl
substituents are placed at the 2 and 7 positions of the fluorenyl moiety of the ligand
(η
5 -C 5 H 4 -μ-CPh 2 -η
5 -2,7-dit
Bu-C 13 H 6 )ZrCl 2 /MAO [131–136] does not show any
marked improvement in the enantioselectivity or stereoselectivity of the resulting
syndiospecific catalyst or in the stereoregularity of its s-PP.
72
A. Razavi