

94
2 The Interaction of Electromagnetic Waves with Water
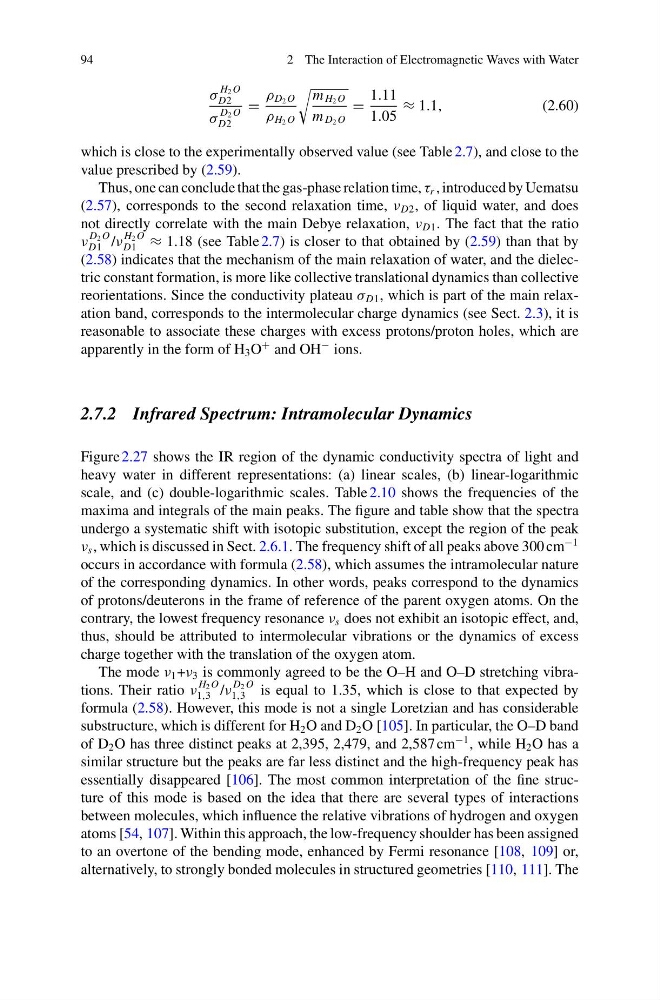
σ
H 2 O
D2
σ
D 2 O
D2
=
ρ D 2 O
ρ H 2 O
m H 2 O
m D 2 O
=
1.11
1.05
≈ 1.1,
(2.60)
which is close to the experimentally observed value (see Table 2.7), and close to the
value prescribed by (2.59).
Thus, one can conclude that the gas-phase relation time, τ r , introduced by Uematsu
(2.57), corresponds to the second relaxation time, ν D2 , of liquid water, and does
not directly correlate with the main Debye relaxation, ν D1 . The fact that the ratio
ν
D 2 O
D1 /ν
H 2 O
D1 ≈ 1.18 (see Table 2.7) is closer to that obtained by (2.59) than that by
(2.58) indicates that the mechanism of the main relaxation of water, and the dielectric constant formation, is more like collective translational dynamics than collective
reorientations. Since the conductivity plateau σ D1 , which is part of the main relaxation band, corresponds to the intermolecular charge dynamics (see Sect. 2.3), it is
reasonable to associate these charges with excess protons/proton holes, which are
apparently in the form of H 3 O
+ and OH
− ions.
2.7.2 Infrared Spectrum: Intramolecular Dynamics
Figure 2.27 shows the IR region of the dynamic conductivity spectra of light and
heavy water in different representations: (a) linear scales, (b) linear-logarithmic
scale, and (c) double-logarithmic scales. Table 2.10 shows the frequencies of the
maxima and integrals of the main peaks. The figure and table show that the spectra
undergo a systematic shift with isotopic substitution, except the region of the peak
ν s , which is discussed in Sect. 2.6.1. The frequency shift of all peaks above 300 cm
−1
occurs in accordance with formula (2.58), which assumes the intramolecular nature
of the corresponding dynamics. In other words, peaks correspond to the dynamics
of protons/deuterons in the frame of reference of the parent oxygen atoms. On the
contrary, the lowest frequency resonance ν s does not exhibit an isotopic effect, and,
thus, should be attributed to intermolecular vibrations or the dynamics of excess
charge together with the translation of the oxygen atom.
The mode ν 1 +ν 3 is commonly agreed to be the O–H and O–D stretching vibrations. Their ratio ν
H 2 O
1,3 /ν
D 2 O
1,3 is equal to 1.35, which is close to that expected by
formula (2.58). However, this mode is not a single Loretzian and has considerable
substructure, which is different for H 2 O and D 2 O [105]. In particular, the O–D band
of D 2 O has three distinct peaks at 2,395, 2,479, and 2,587 cm
−1 , while H 2 O has a
similar structure but the peaks are far less distinct and the high-frequency peak has
essentially disappeared [106]. The most common interpretation of the fine structure of this mode is based on the idea that there are several types of interactions
between molecules, which influence the relative vibrations of hydrogen and oxygen
atoms [54, 107]. Within this approach, the low-frequency shoulder has been assigned
to an overtone of the bending mode, enhanced by Fermi resonance [108, 109] or,
alternatively, to strongly bonded molecules in structured geometries [110, 111]. The