

3.2 Calculation Methods for Simulating Dopaquinone Conversion
57
analysis [28]. Considering aqueous phase reactions, we used PCM to describe the
solvent–solute interaction [29, 30].
As a descriptor of nucleophilicity, the condensed-to-atom Fukui indices (the
derivatives of the electron density with respect to the total number of electrons,
namely the normalized local softness of the electronic system) were calculated using
the finite difference approximation [31, 32]. In this approximation, the N + 1 and N
− 1 electron systems were calculated using the structure of the N electron system, and
then the atomic charges were determined by the natural population orbital analysis.
To obtain the activation barriers, one-dimensionally projected potential energy
curves were calculated along the direction in which the bond length increases with
a step size increment of 0.05 or 0.10 Å. During the calculations, all the degrees
of freedom except for the one specifically chosen to be frozen (as specified by the
structure of the reaction) were allowed to relax. To find the transition states of the
reactions, we used the synchronous transit and quasi-Newton (STQN) method [33].
Note that the transition state structures and the activation energies obtained from the
calculated potential energy curves and from the STQN method were almost identical.
3.3 Competition Between Cyclization and Thiol
Binding—Comparison Between Dopaquinone
and Rhododendrol Quinone
As described in Sect. 3.1, o-quinones competitively undergo cyclization and thiol
binding even though their active sites do not overlap. Here, we considered metastable
species before and after cyclization to discuss the change of the binding ability to
thiols. As a model of thiols, the simplest structure methane thiolate ion (SCH 3
− )
was chosen. For the comparison of o-quinone structures, we chose dopaquinone and
RD-quinone.
First, we investigated the change in the electronic state and the total energy
when SCH 3
− ion binds with dopaquinone. As shown in Fig. 3.4, through the C–
S bond formation, a charge transfer of approximately one electron occurred into
dopaquinone, which occupied the LUMO of dopaquinone. With this, the change in
the total energy of this bond formation would be mainly determined by the LUMO
level of dopaquinone. The binding energy is calculated as
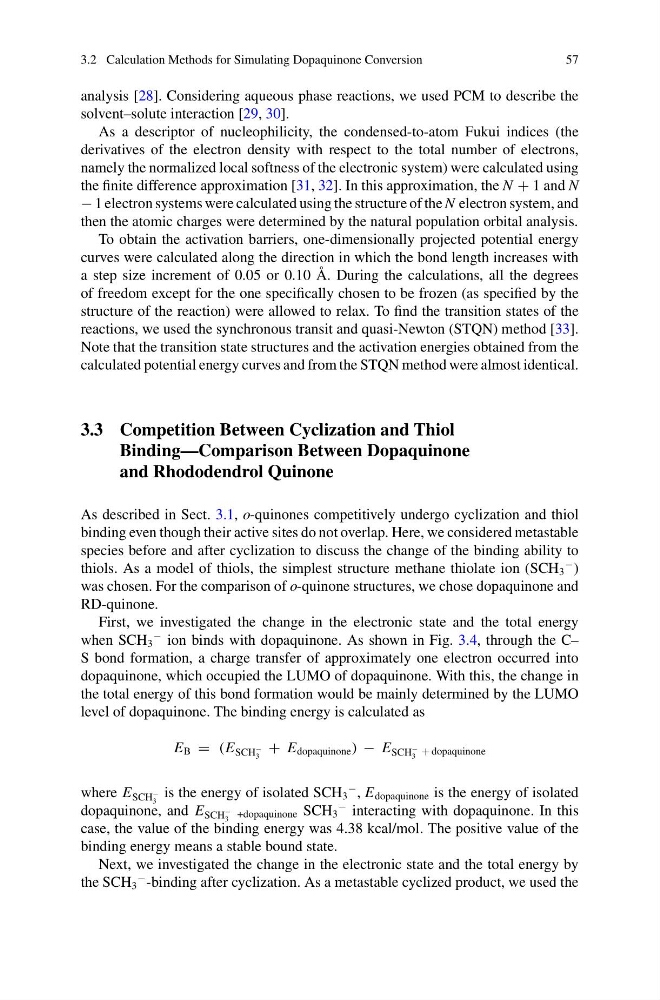
E B = (E SCH
−
3
+ E dopaquinone ) − E SCH
−
3
+ dopaquinone
where E SCH
−
3
is the energy of isolated SCH 3
− , E dopaquinone is the energy of isolated
dopaquinone, and E SCH
−
3
+dopaquinone SCH 3
− interacting with dopaquinone. In this
case, the value of the binding energy was 4.38 kcal/mol. The positive value of the
binding energy means a stable bound state.
Next, we investigated the change in the electronic state and the total energy by
the SCH 3
− -binding after cyclization. As a metastable cyclized product, we used the