

188
S. Sakkiah et al.
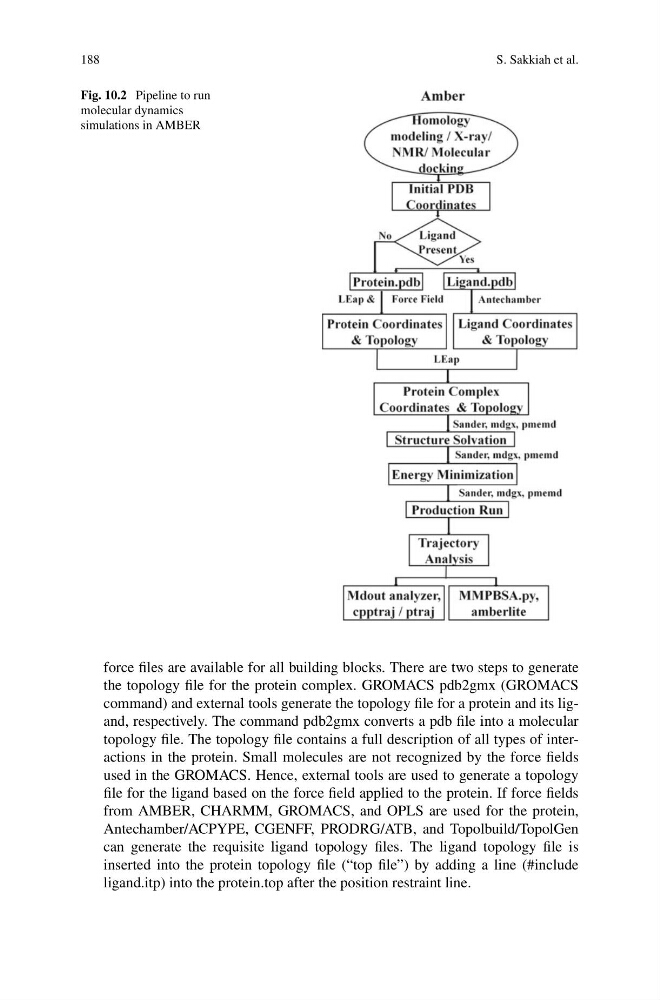
Fig. 10.2 Pipeline to run
molecular dynamics
simulations in AMBER
force files are available for all building blocks. There are two steps to generate
the topology file for the protein complex. GROMACS pdb2gmx (GROMACS
command) and external tools generate the topology file for a protein and its ligand, respectively. The command pdb2gmx converts a pdb file into a molecular
topology file. The topology file contains a full description of all types of interactions in the protein. Small molecules are not recognized by the force fields
used in the GROMACS. Hence, external tools are used to generate a topology
file for the ligand based on the force field applied to the protein. If force fields
from AMBER, CHARMM, GROMACS, and OPLS are used for the protein,
Antechamber/ACPYPE, CGENFF, PRODRG/ATB, and Topolbuild/TopolGen
can generate the requisite ligand topology files. The ligand topology file is
inserted into the protein topology file (“top file”) by adding a line (#include
ligand.itp) into the protein.top after the position restraint line.