

10 Applications of Molecular Dynamics Simulations …
187
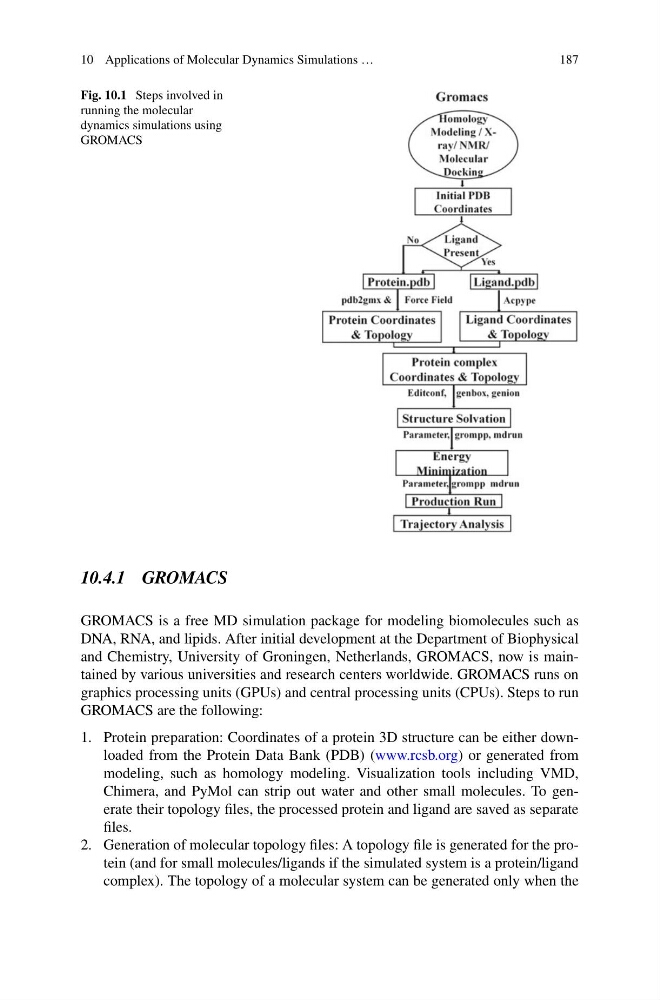
Fig. 10.1 Steps involved in
running the molecular
dynamics simulations using
GROMACS
10.4.1 GROMACS
GROMACS is a free MD simulation package for modeling biomolecules such as
DNA, RNA, and lipids. After initial development at the Department of Biophysical
and Chemistry, University of Groningen, Netherlands, GROMACS, now is maintained by various universities and research centers worldwide. GROMACS runs on
graphics processing units (GPUs) and central processing units (CPUs). Steps to run
GROMACS are the following:
1. Protein preparation: Coordinates of a protein 3D structure can be either downloaded from the Protein Data Bank (PDB) (www.rcsb.org) or generated from
modeling, such as homology modeling. Visualization tools including VMD,
Chimera, and PyMol can strip out water and other small molecules. To generate their topology files, the processed protein and ligand are saved as separate
files.
2. Generation of molecular topology files: A topology file is generated for the protein (and for small molecules/ligands if the simulated system is a protein/ligand
complex). The topology of a molecular system can be generated only when the