

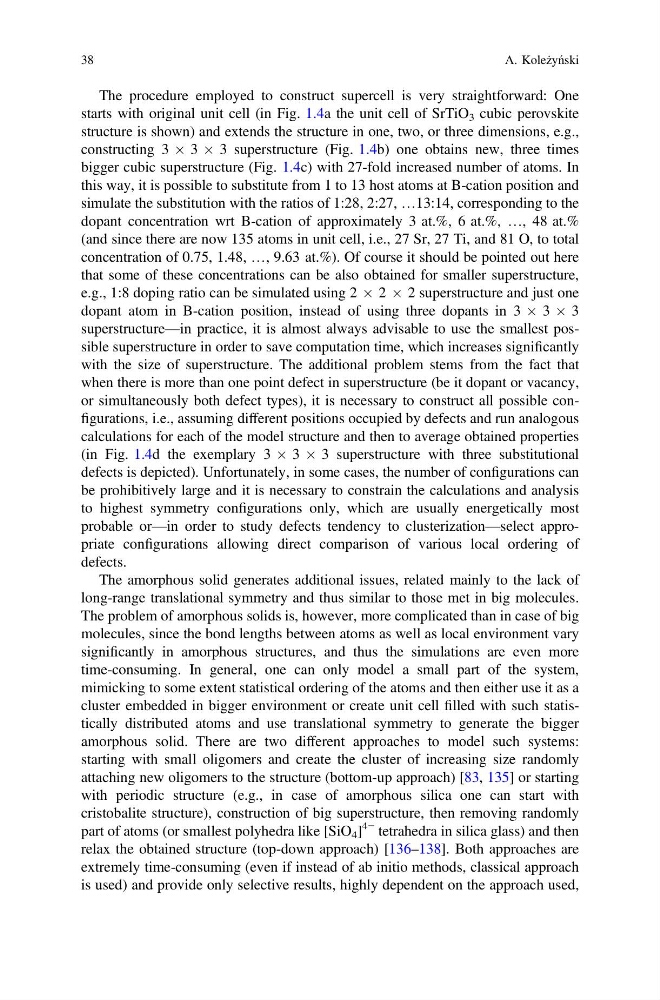
The procedure employed to construct supercell is very straightforward: One
starts with original unit cell (in Fig. 1.4a the unit cell of SrTiO 3 cubic perovskite
structure is shown) and extends the structure in one, two, or three dimensions, e.g.,
constructing 3 Â 3 Â 3 superstructure (Fig. 1.4b) one obtains new, three times
bigger cubic superstructure (Fig. 1.4c) with 27-fold increased number of atoms. In
this way, it is possible to substitute from 1 to 13 host atoms at B-cation position and
simulate the substitution with the ratios of 1:28, 2:27, …13:14, corresponding to the
dopant concentration wrt B-cation of approximately 3 at.%, 6 at.%, …, 48 at.%
(and since there are now 135 atoms in unit cell, i.e., 27 Sr, 27 Ti, and 81 O, to total
concentration of 0.75, 1.48, …, 9.63 at.%). Of course it should be pointed out here
that some of these concentrations can be also obtained for smaller superstructure,
e.g., 1:8 doping ratio can be simulated using 2 Â 2 Â 2 superstructure and just one
dopant atom in B-cation position, instead of using three dopants in 3 Â 3 Â 3
superstructure—in practice, it is almost always advisable to use the smallest possible superstructure in order to save computation time, which increases significantly
with the size of superstructure. The additional problem stems from the fact that
when there is more than one point defect in superstructure (be it dopant or vacancy,
or simultaneously both defect types), it is necessary to construct all possible configurations, i.e., assuming different positions occupied by defects and run analogous
calculations for each of the model structure and then to average obtained properties
(in Fig. 1.4d the exemplary 3 Â 3 Â 3 superstructure with three substitutional
defects is depicted). Unfortunately, in some cases, the number of configurations can
be prohibitively large and it is necessary to constrain the calculations and analysis
to highest symmetry configurations only, which are usually energetically most
probable or—in order to study defects tendency to clusterization—select appropriate configurations allowing direct comparison of various local ordering of
defects.
The amorphous solid generates additional issues, related mainly to the lack of
long-range translational symmetry and thus similar to those met in big molecules.
The problem of amorphous solids is, however, more complicated than in case of big
molecules, since the bond lengths between atoms as well as local environment vary
significantly in amorphous structures, and thus the simulations are even more
time-consuming. In general, one can only model a small part of the system,
mimicking to some extent statistical ordering of the atoms and then either use it as a
cluster embedded in bigger environment or create unit cell filled with such statistically distributed atoms and use translational symmetry to generate the bigger
amorphous solid. There are two different approaches to model such systems:
starting with small oligomers and create the cluster of increasing size randomly
attaching new oligomers to the structure (bottom-up approach) [83, 135] or starting
with periodic structure (e.g., in case of amorphous silica one can start with
cristobalite structure), construction of big superstructure, then removing randomly
part of atoms (or smallest polyhedra like [SiO 4 ]
4− tetrahedra in silica glass) and then
relax the obtained structure (top-down approach) [136–138]. Both approaches are
extremely time-consuming (even if instead of ab initio methods, classical approach
is used) and provide only selective results, highly dependent on the approach used,
38
A. Koleżyński