

172
8 Hetero-Coordinated Interfaces
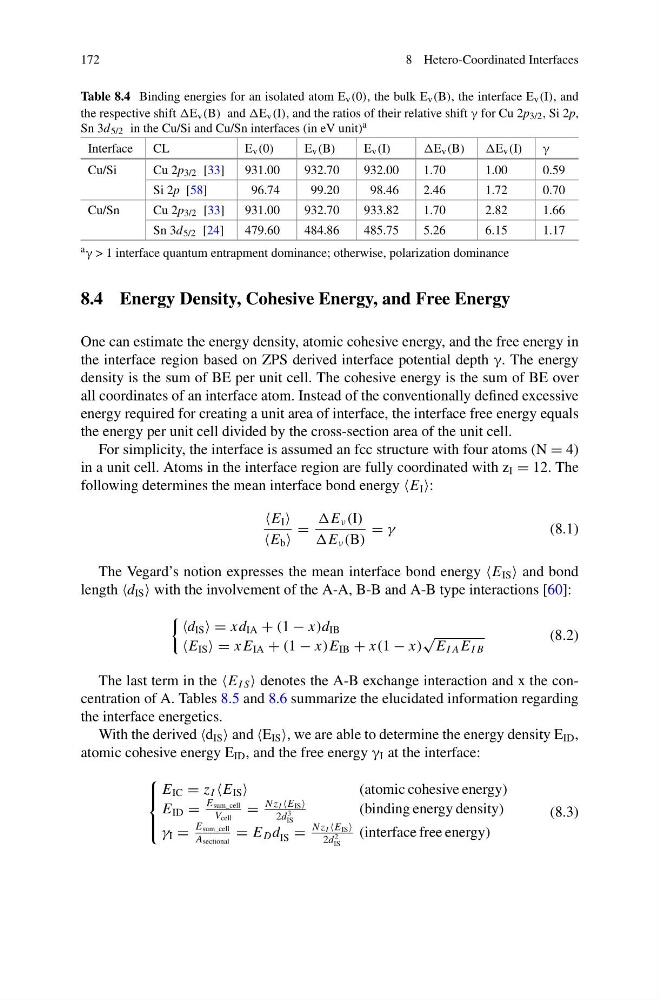
Table 8.4 Binding energies for an isolated atom E v (0), the bulk E v (B), the interface E v (I), and
the respective shift E v (B) and E v (I), and the ratios of their relative shift γ for Cu 2p 3/2 , Si 2p,
Sn 3d 5/2 in the Cu/Si and Cu/Sn interfaces (in eV unit) a
Interface
CL
E v (0)
E v (B)
E v (I)
E v (B)
E v (I)
γ
Cu/Si
Cu 2p 3/2 [33]
931.00
932.70
932.00
1.70
1.00
0.59
Si 2p [58]
96.74
99.20
98.46
2.46
1.72
0.70
Cu/Sn
Cu 2p 3/2 [33]
931.00
932.70
933.82
1.70
2.82
1.66
Sn 3d 5/2 [24]
479.60
484.86
485.75
5.26
6.15
1.17
a γ > 1 interface quantum entrapment dominance; otherwise, polarization dominance
8.4 Energy Density, Cohesive Energy, and Free Energy
One can estimate the energy density, atomic cohesive energy, and the free energy in
the interface region based on ZPS derived interface potential depth γ. The energy
density is the sum of BE per unit cell. The cohesive energy is the sum of BE over
all coordinates of an interface atom. Instead of the conventionally defined excessive
energy required for creating a unit area of interface, the interface free energy equals
the energy per unit cell divided by the cross-section area of the unit cell.
For simplicity, the interface is assumed an fcc structure with four atoms (N = 4)
in a unit cell. Atoms in the interface region are fully coordinated with z I = 12. The
following determines the mean interface bond energy E I :
E I
E b
=
E ν (I)
E ν (B)
= γ
(8.1)
The Vegard’s notion expresses the mean interface bond energy E IS and bond
length d IS with the involvement of the A-A, B-B and A-B type interactions [60]:
d IS = xd IA + (1 − x)d IB
E IS = x E IA + (1 − x)E IB + x(1 − x)
√
E I A E I B
(8.2)
The last term in the E I S denotes the A-B exchange interaction and x the concentration of A. Tables 8.5 and 8.6 summarize the elucidated information regarding
the interface energetics.
With the derived d IS and E IS , we are able to determine the energy density E ID ,
atomic cohesive energy E ID , and the free energy γ I at the interface:
⎧
⎪ ⎨
⎪ ⎩
E IC = z I E IS
(atomic cohesive energy)
E ID =
E sum_cell
V cell
=
N z I E IS
2d
3
IS
(binding energy density)
γ I =
E sum_cell
A sectional
= E D d IS =
N z I E IS
2d
2
IS
(interface free energy)
(8.3)