

48
Download the latest UCSF Chimera release for your operating system form https://www.cgl.ucsf.edu/chimera/. Follow
the installation instructions.
3. Swiss Param:
Visit the Swiss Param server website in order to create your
molecule topologies at http://www.swissparam.ch/ (further
directions are in Subheading 3).
4. Gromacs:
Install the latest Gromacs release at your Linux workstation
or check for the available modules at your HPC server, http://
www.gromacs.org/.
5. VMD:
Download and install the latest VMD release for your operating
system, https://www.ks.uiuc.edu/Development/Download/
download.cgi?PackageName=VMD.
3 Methods
All the MD simulations were carried out using the Gromacs/2018.1
simulation package and the workflow below is adjusted for this
specific software [8]. Similar workflows can be used for different
Reaction Coordinate
energy
function for
each
window
configuration
window
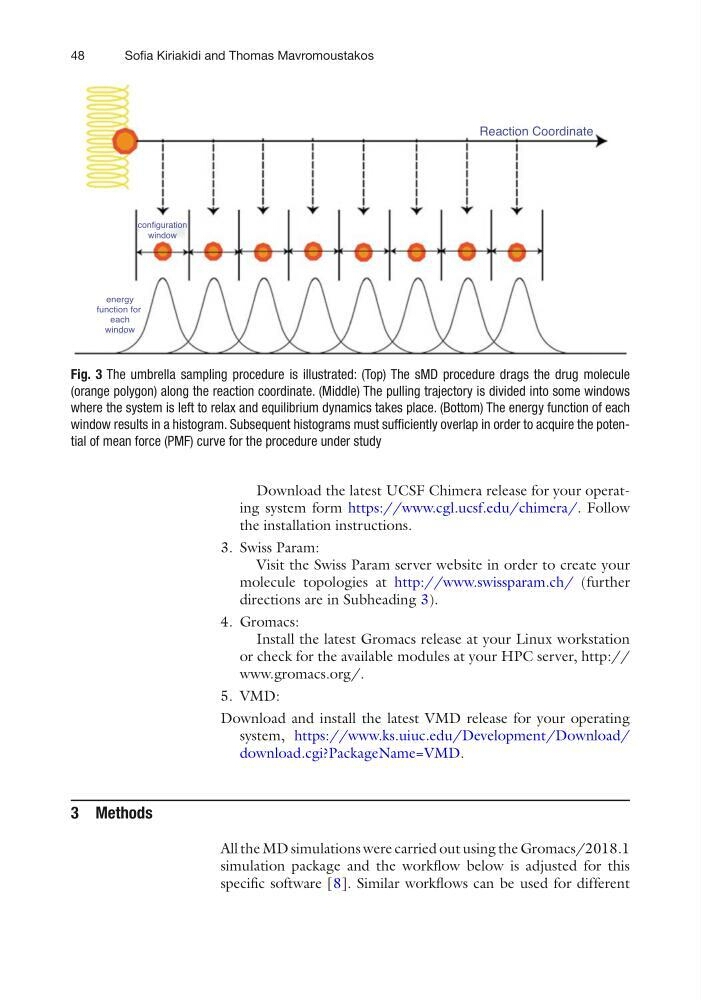
Fig. 3 The umbrella sampling procedure is illustrated: (Top) The sMD procedure drags the drug molecule
(orange polygon) along the reaction coordinate. (Middle) The pulling trajectory is divided into some windows
where the system is left to relax and equilibrium dynamics takes place. (Bottom) The energy function of each
window results in a histogram. Subsequent histograms must sufficiently overlap in order to acquire the potential of mean force (PMF) curve for the procedure under study
Sofia Kiriakidi and Thomas Mavromoustakos