

7.2.9 VCF
The VCF (Variant Call Format) contains information about genetic variants found at
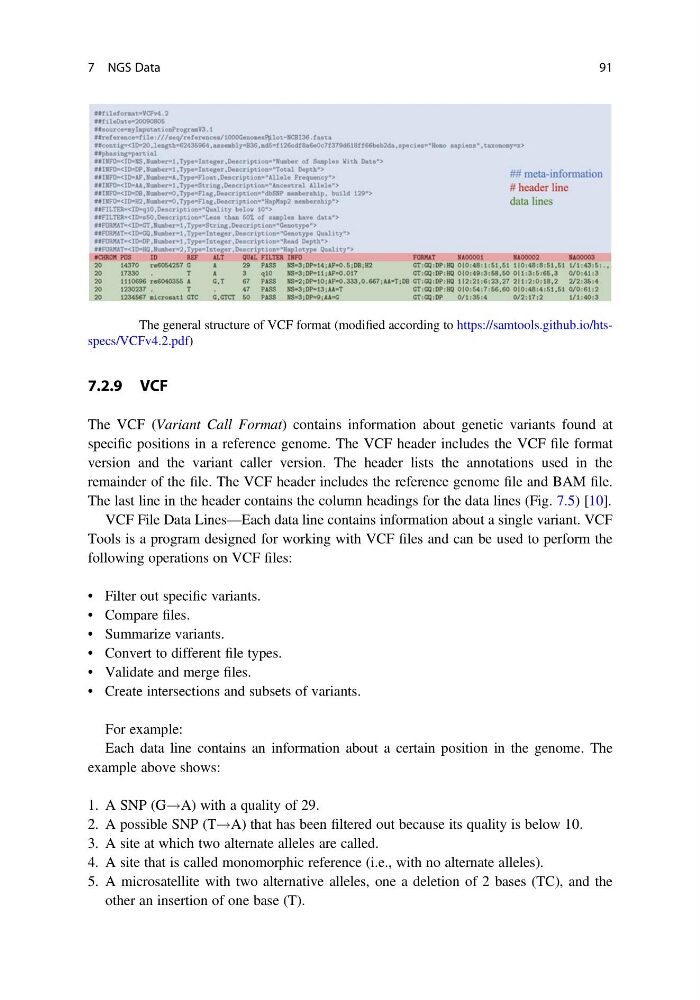
specific positions in a reference genome. The VCF header includes the VCF file format
version and the variant caller version. The header lists the annotations used in the
remainder of the file. The VCF header includes the reference genome file and BAM file.
The last line in the header contains the column headings for the data lines (Fig. 7.5) [10].
VCF File Data Lines—Each data line contains information about a single variant. VCF
Tools is a program designed for working with VCF files and can be used to perform the
following operations on VCF files:
• Filter out specific variants.
• Compare files.
• Summarize variants.
• Convert to different file types.
• Validate and merge files.
• Create intersections and subsets of variants.
For example:
Each data line contains an information about a certain position in the genome. The
example above shows:
1. A SNP (G!A) with a quality of 29.
2. A possible SNP (T!A) that has been filtered out because its quality is below 10.
3. A site at which two alternate alleles are called.
4. A site that is called monomorphic reference (i.e., with no alternate alleles).
5. A microsatellite with two alternative alleles, one a deletion of 2 bases (TC), and the
other an insertion of one base (T).
Fig. 7.5 The general structure of VCF format (modified according to https://samtools.github.io/htsspecs/VCFv4.2.pdf)
7 NGS Data
91