

considerable charge transfer upon excitation, as they include groups that have
different electron affinities (EA) and ionisation energies (IE) in the gas phase and
are coupled via conjugated parts. In so-called charge-transfer (CT) systems, an
electron donating group may transfer electron density to an acceptor group upon
excitation. The charge localisation in the ground state and the excitation-induced
charge transfer can considerably depend on the geometry and the employed QM
method. This can be rationalised for a given system in terms of the involved
resonance structures. The contribution of these resonances to the ground and
excited state determines both charge localisation and bond length alternation of
the minimum energy structure in the respective electronic state. The balance
between different valence bond structures often strongly depends on the description
of electron correlation by the QM method, and, in the case of time-dependent
density functional theory (TDDFT), on the employed density functional. In the
following, we will discuss the advantages and typical shortcomings of the most
commonly used QM methods and the resulting performance in the description of
perturbations of the kind (1)–(4). Moreover, the description of photochemical
properties requires an analysis of the excited-state potential energy surface (PES).
Already the qualitatively correct description of the excited-state PES, i.e., its
topology in terms of minima, conical intersections and transition states, represents
a major challenge for state-of-the-art quantum chemical methods. This will be
discussed in Sects. 4.2 and 4.4.
4.2.1 TDDFT
The extension of density functional theory (DFT) to the time domain [3] provides
access to excited-state properties. If the ground state is perturbed by an external
field of the form v ext (t) ¼ xδ(t À t 0 ) (small instantaneous kick), the perturbed
density can be propagated in time following the time-dependent Kohn-Sham
equations. The absorption spectrum can then be obtained from the Fourier
transformed dipole moment. This approach is useful to calculate the spectrum
within a large energy window, e.g., beyond the ionisation threshold, and for large
systems (high density of states), where linear scaling of cpu time vs. system size can
easily be achieved [4]. The more common approach is to solve analytically the
linear-response problem in a basis of molecular orbitals to obtain a limited number
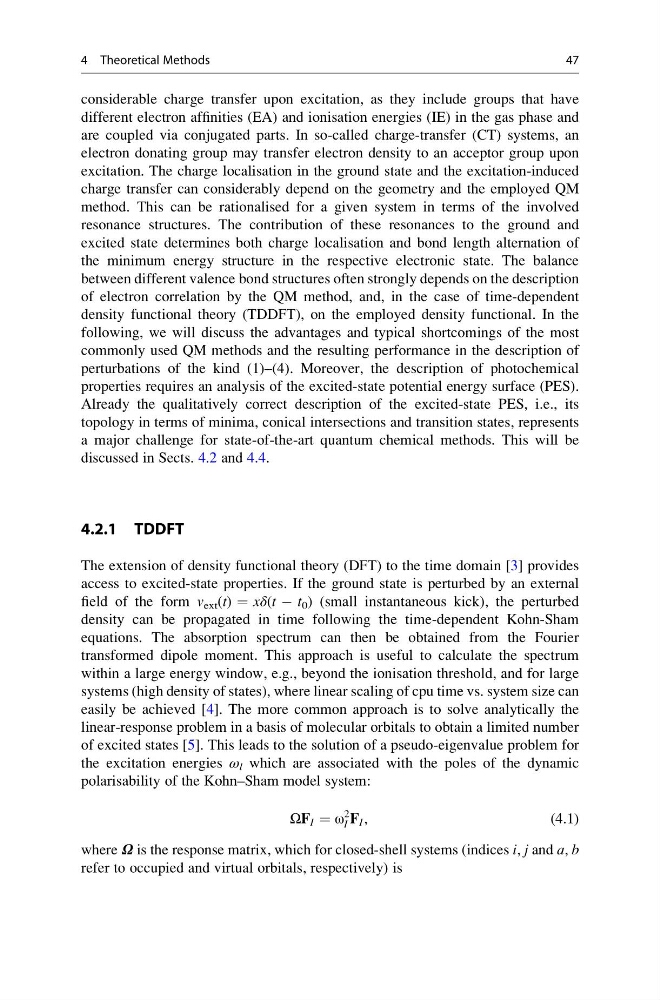
of excited states [5]. This leads to the solution of a pseudo-eigenvalue problem for
the excitation energies ω l which are associated with the poles of the dynamic
polarisability of the Kohn–Sham model system:
ΩF I ¼ ω
2
I F I ,
(4.1)
where Ω is the response matrix, which for closed-shell systems (indices i, j and a, b
refer to occupied and virtual orbitals, respectively) is
4 Theoretical Methods
47