

196
6 Magnetism and Conduction
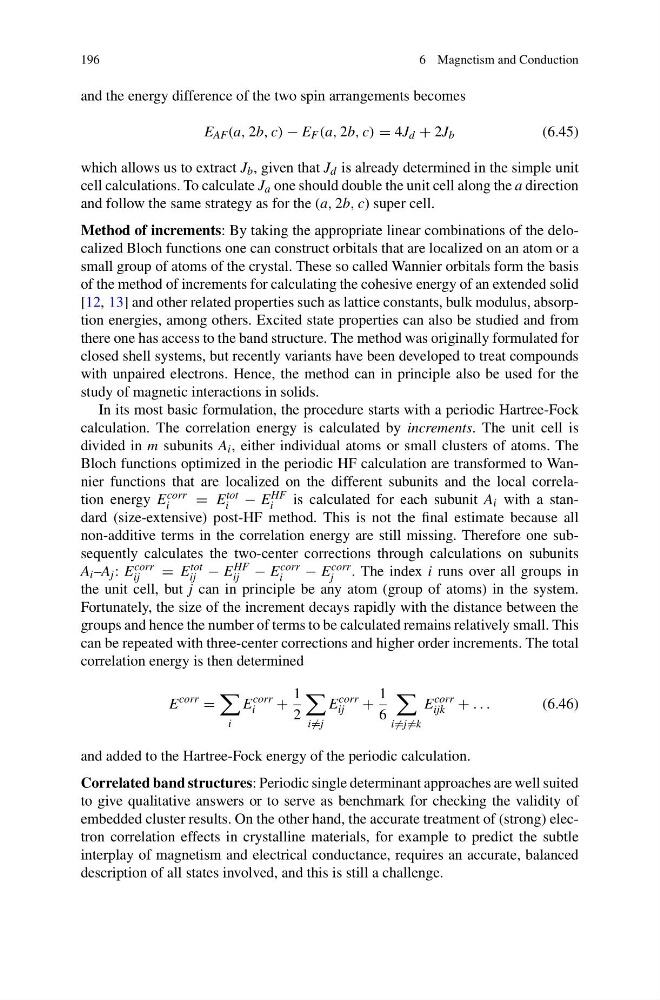
and the energy difference of the two spin arrangements becomes
E AF (a, 2b, c) − E F (a, 2b, c) = 4J d + 2J b
(6.45)
which allows us to extract J b , given that J d is already determined in the simple unit
cell calculations. To calculate J a one should double the unit cell along the a direction
and follow the same strategy as for the (a, 2b, c) super cell.
Method of increments: By taking the appropriate linear combinations of the delocalized Bloch functions one can construct orbitals that are localized on an atom or a
small group of atoms of the crystal. These so called Wannier orbitals form the basis
of the method of increments for calculating the cohesive energy of an extended solid
[12, 13] and other related properties such as lattice constants, bulk modulus, absorption energies, among others. Excited state properties can also be studied and from
there one has access to the band structure. The method was originally formulated for
closed shell systems, but recently variants have been developed to treat compounds
with unpaired electrons. Hence, the method can in principle also be used for the
study of magnetic interactions in solids.
In its most basic formulation, the procedure starts with a periodic Hartree-Fock
calculation. The correlation energy is calculated by increments. The unit cell is
divided in m subunits A i , either individual atoms or small clusters of atoms. The
Bloch functions optimized in the periodic HF calculation are transformed to Wannier functions that are localized on the different subunits and the local correlation energy E corr
i
= E tot
i − E HF
i
is calculated for each subunit A i with a standard (size-extensive) post-HF method. This is not the final estimate because all
non-additive terms in the correlation energy are still missing. Therefore one subsequently calculates the two-center corrections through calculations on subunits
A i –A j : E corr
ij
= E tot
ij − E HF
ij − E corr
i
− E corr
j
. The index i runs over all groups in
the unit cell, but j can in principle be any atom (group of atoms) in the system.
Fortunately, the size of the increment decays rapidly with the distance between the
groups and hence the number of terms to be calculated remains relatively small. This
can be repeated with three-center corrections and higher order increments. The total
correlation energy is then determined
E
corr =
i
E
corr
i
+
1
2
i =j
E
corr
ij
+
1
6
i =j =k
E
corr
ijk + ...
(6.46)
and added to the Hartree-Fock energy of the periodic calculation.
Correlated band structures: Periodic single determinant approaches are well suited
to give qualitative answers or to serve as benchmark for checking the validity of
embedded cluster results. On the other hand, the accurate treatment of (strong) electron correlation effects in crystalline materials, for example to predict the subtle
interplay of magnetism and electrical conductance, requires an accurate, balanced
description of all states involved, and this is still a challenge.