

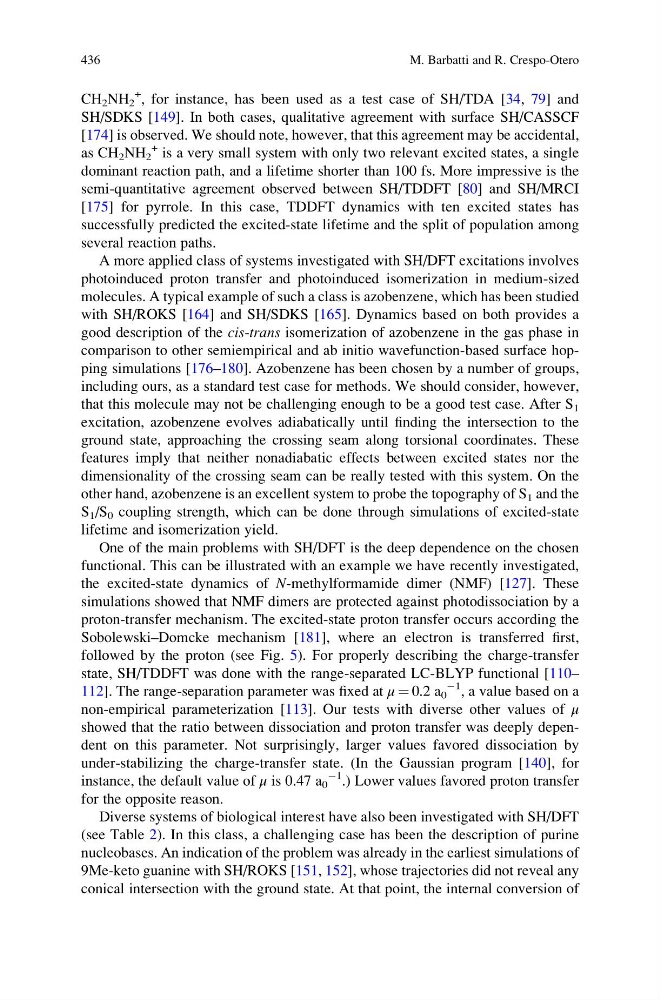
CH 2 NH 2
+
, for instance, has been used as a test case of SH/TDA [34, 79] and
SH/SDKS [149]. In both cases, qualitative agreement with surface SH/CASSCF
[174] is observed. We should note, however, that this agreement may be accidental,
as CH 2 NH 2
+ is a very small system with only two relevant excited states, a single
dominant reaction path, and a lifetime shorter than 100 fs. More impressive is the
semi-quantitative agreement observed between SH/TDDFT [80] and SH/MRCI
[175] for pyrrole. In this case, TDDFT dynamics with ten excited states has
successfully predicted the excited-state lifetime and the split of population among
several reaction paths.
A more applied class of systems investigated with SH/DFT excitations involves
photoinduced proton transfer and photoinduced isomerization in medium-sized
molecules. A typical example of such a class is azobenzene, which has been studied
with SH/ROKS [164] and SH/SDKS [165]. Dynamics based on both provides a
good description of the cis-trans isomerization of azobenzene in the gas phase in
comparison to other semiempirical and ab initio wavefunction-based surface hopping simulations [176–180]. Azobenzene has been chosen by a number of groups,
including ours, as a standard test case for methods. We should consider, however,
that this molecule may not be challenging enough to be a good test case. After S 1
excitation, azobenzene evolves adiabatically until finding the intersection to the
ground state, approaching the crossing seam along torsional coordinates. These
features imply that neither nonadiabatic effects between excited states nor the
dimensionality of the crossing seam can be really tested with this system. On the
other hand, azobenzene is an excellent system to probe the topography of S 1 and the
S 1 /S 0 coupling strength, which can be done through simulations of excited-state
lifetime and isomerization yield.
One of the main problems with SH/DFT is the deep dependence on the chosen
functional. This can be illustrated with an example we have recently investigated,
the excited-state dynamics of N-methylformamide dimer (NMF) [127]. These
simulations showed that NMF dimers are protected against photodissociation by a
proton-transfer mechanism. The excited-state proton transfer occurs according the
Sobolewski–Domcke mechanism [181], where an electron is transferred first,
followed by the proton (see Fig. 5). For properly describing the charge-transfer
state, SH/TDDFT was done with the range-separated LC-BLYP functional [110–
112]. The range-separation parameter was fixed at μ ¼ 0.2 a 0
À1 , a value based on a
non-empirical parameterization [113]. Our tests with diverse other values of μ
showed that the ratio between dissociation and proton transfer was deeply dependent on this parameter. Not surprisingly, larger values favored dissociation by
under-stabilizing the charge-transfer state. (In the Gaussian program [140], for
instance, the default value of μ is 0.47 a 0
À1 .) Lower values favored proton transfer
for the opposite reason.
Diverse systems of biological interest have also been investigated with SH/DFT
(see Table 2). In this class, a challenging case has been the description of purine
nucleobases. An indication of the problem was already in the earliest simulations of
9Me-keto guanine with SH/ROKS [151, 152], whose trajectories did not reveal any
conical intersection with the ground state. At that point, the internal conversion of
436
M. Barbatti and R. Crespo-Otero