

Nano-Porous Graphene as Free-Standing Membranes
55
Table 1 summarizes the configurations and parameters of MD results on pristine and
functionalized nano-porous graphene for water desalination.
MD simulations are ideal to study nano-porous graphene system for water desalination since they compute the evolution of a system of atoms from an original configuration and under a set of constraints [111]. In these simulations, the forces between
atoms at each time step are calculated, then, the positions of all the atoms at the
following time step is updated using Newton’s equations of motion [99]. Depending
on the choice of force field, the performance of relatively large molecular systems
(10
2
−10
9 atoms) can be explored over physically meaningful time scales (typically
between 1 ns and 1 µs) [99]. With this introduction, MD simulations allow us to probe
the kinetics and thermodynamics of desalination while accounting for the physics of
water, ions, and graphene layers with high accuracy [99].
MD simulation is usually performed with the temperature maintained at 300 K
by using a Nose-Hoover thermostat. However, there are a few reports that studied
water desalination parameters of functionalized nano-porous graphene membrane at
a range of temperatures 275–343 K [17]. Periodic boundary conditions are employed
to maintain a continuous 2D membrane. In some simulations, the functional groups
were usually pointed toward the center of the pore due to steric constraints [104].
Thus, the diameter of the pore changed depending on the size of the functional groups
grafted to it. The diameter of the functionalized pore was calculated as the distance
between the ends of two opposite functional groups. In each functionalized pore, a
specific number of functional groups were spread uniformly around the rim of the
pore [17, 104]. In other works, the size of each pore was measured by plotting atoms as
van der Waals spheres and calculating the amount of contiguous area not obstructed
by any atomic representations [98]. Pore diameters were obtained from the open
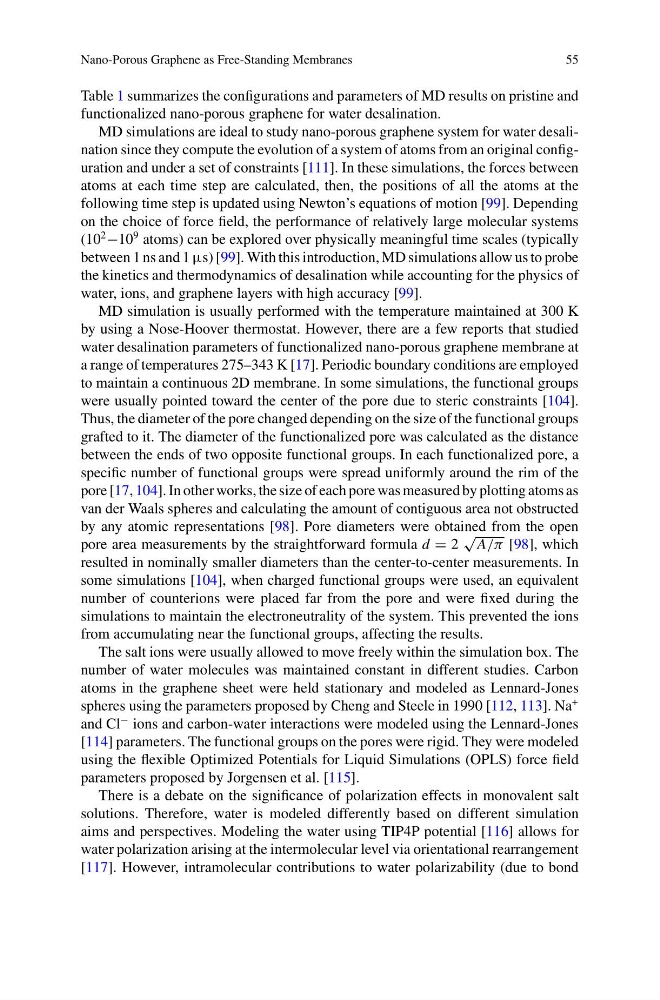
pore area measurements by the straightforward formula d = 2
√
A/π [98], which
resulted in nominally smaller diameters than the center-to-center measurements. In
some simulations [104], when charged functional groups were used, an equivalent
number of counterions were placed far from the pore and were fixed during the
simulations to maintain the electroneutrality of the system. This prevented the ions
from accumulating near the functional groups, affecting the results.
The salt ions were usually allowed to move freely within the simulation box. The
number of water molecules was maintained constant in different studies. Carbon
atoms in the graphene sheet were held stationary and modeled as Lennard-Jones
spheres using the parameters proposed by Cheng and Steele in 1990 [112, 113]. Na
+
and Cl
− ions and carbon-water interactions were modeled using the Lennard-Jones
[114] parameters. The functional groups on the pores were rigid. They were modeled
using the flexible Optimized Potentials for Liquid Simulations (OPLS) force field
parameters proposed by Jorgensen et al. [115].
There is a debate on the significance of polarization effects in monovalent salt
solutions. Therefore, water is modeled differently based on different simulation
aims and perspectives. Modeling the water using TIP4P potential [116] allows for
water polarization arising at the intermolecular level via orientational rearrangement
[117]. However, intramolecular contributions to water polarizability (due to bond