

590
A. V. Luzanov
which makes sense only for computations of G
QCTB
0
by (2). Here δ is an orbital
splitting parameter by which different orbital for different spins are introduced into
the TB scheme, thus taking into implicit account the π -electron correlation effects.
A further improvement is attained by combining QCTB and RHF models. The
approach is suggested in [21] and termed the extended quasi-correlated (EQC)
method. The preliminary results were given in [19] in terms of the quasi-correlated
long-range interaction (QCLRI) scheme which is identical to EQC from [21]. In this
case we construct the intermediate Fock matrix f [1] (also shifted by the Fermi energy).
This matrix is normally computed by using the TB density matrix (bond-order matrix)
P. Hence, we do not invoke any self-consistency procedure. The obtained intermediate Fockian f [1] is then renormalized [see (25)] and used straightforwardly in
computing G
EQC
0
by (2) with applying a formal Hamiltonian similar to (5):
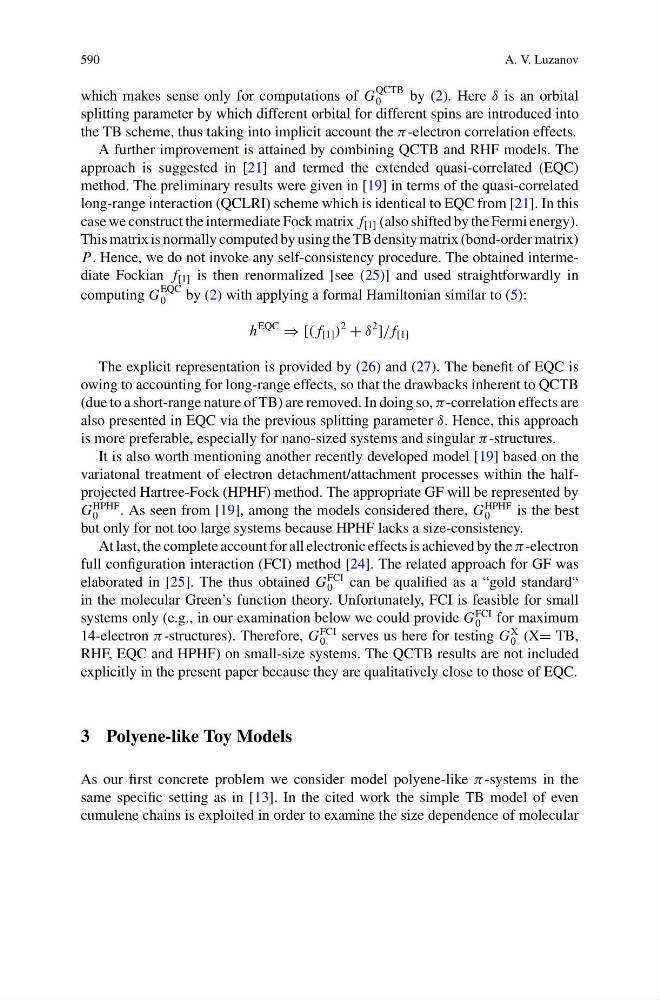
h
EQC
⇒ [( f [1] )
2
+ δ
2
]/ f [1]
The explicit representation is provided by (26) and (27). The benefit of EQC is
owing to accounting for long-range effects, so that the drawbacks inherent to QCTB
(due to a short-range nature of TB) are removed. In doing so, π -correlation effects are
also presented in EQC via the previous splitting parameter δ. Hence, this approach
is more preferable, especially for nano-sized systems and singular π -structures.
It is also worth mentioning another recently developed model [19] based on the
variatonal treatment of electron detachment/attachment processes within the halfprojected Hartree-Fock (HPHF) method. The appropriate GF will be represented by
G
HPHF
0
. As seen from [19], among the models considered there, G
HPHF
0
is the best
but only for not too large systems because HPHF lacks a size-consistency.
At last, the complete account for all electronic effects is achieved by the π -electron
full configuration interaction (FCI) method [24]. The related approach for GF was
elaborated in [25]. The thus obtained G
FCI
0
can be qualified as a “gold standard“
in the molecular Green’s function theory. Unfortunately, FCI is feasible for small
systems only (e.g., in our examination below we could provide G
FCI
0
for maximum
14-electron π -structures). Therefore, G
FCI
0
serves us here for testing G
X
0 (X= TB,
RHF, EQC and HPHF) on small-size systems. The QCTB results are not included
explicitly in the present paper because they are qualitatively close to those of EQC.
3 Polyene-like Toy Models
As our first concrete problem we consider model polyene-like π -systems in the
same specific setting as in [13]. In the cited work the simple TB model of even
cumulene chains is exploited in order to examine the size dependence of molecular