

5.3 Embarrassingly Parallel Molecular Docking Pipeline
Molecular docking or high-throughput screening has become increasingly important in the context of drug discovery [45]. High-throughput screening may be the
only way to identify correct inhibitors of the specific target. However,
high-throughput drug docking is cost-effective and very fast and could be very
useful for pharmaceutical industry. An attempt has been made to develop a scalable
workflow as shown in Fig. 12, for high-throughput conformational search and
docking on the high-performance computing, Hadoop or cloud-based clusters. The
workflow is divided into two sections. The first section performs conformational
search, and the second section performs the molecular docking. The objective of the
conformation search is to find the most stable conformation of the molecule along
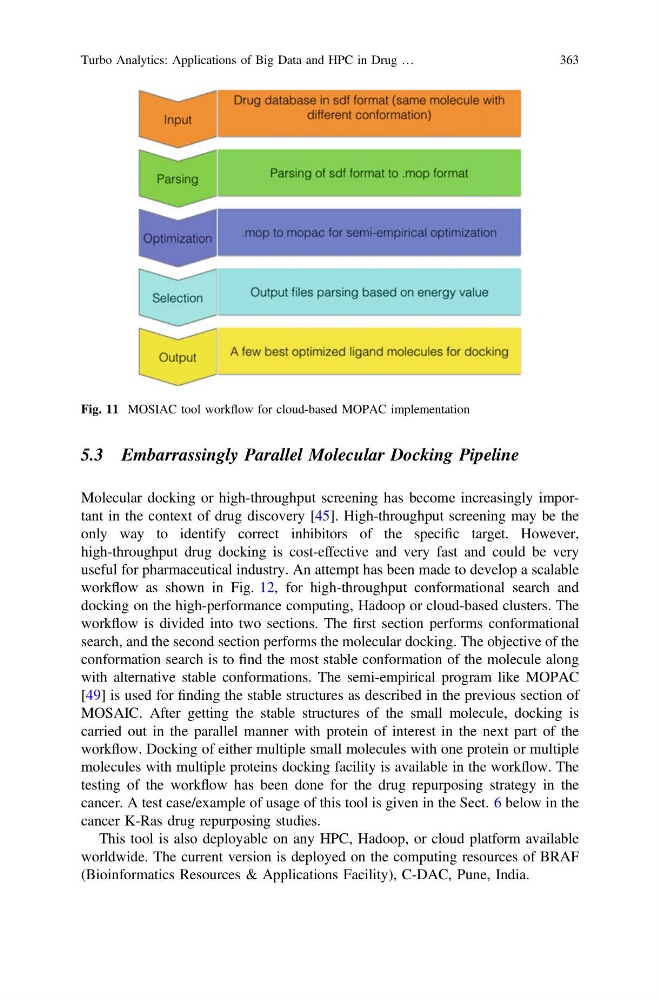
with alternative stable conformations. The semi-empirical program like MOPAC
[49] is used for finding the stable structures as described in the previous section of
MOSAIC. After getting the stable structures of the small molecule, docking is
carried out in the parallel manner with protein of interest in the next part of the
workflow. Docking of either multiple small molecules with one protein or multiple
molecules with multiple proteins docking facility is available in the workflow. The
testing of the workflow has been done for the drug repurposing strategy in the
cancer. A test case/example of usage of this tool is given in the Sect. 6 below in the
cancer K-Ras drug repurposing studies.
This tool is also deployable on any HPC, Hadoop, or cloud platform available
worldwide. The current version is deployed on the computing resources of BRAF
(Bioinformatics Resources & Applications Facility), C-DAC, Pune, India.
Fig. 11 MOSIAC tool workflow for cloud-based MOPAC implementation
Turbo Analytics: Applications of Big Data and HPC in Drug …
363