

190
S. Sakkiah et al.
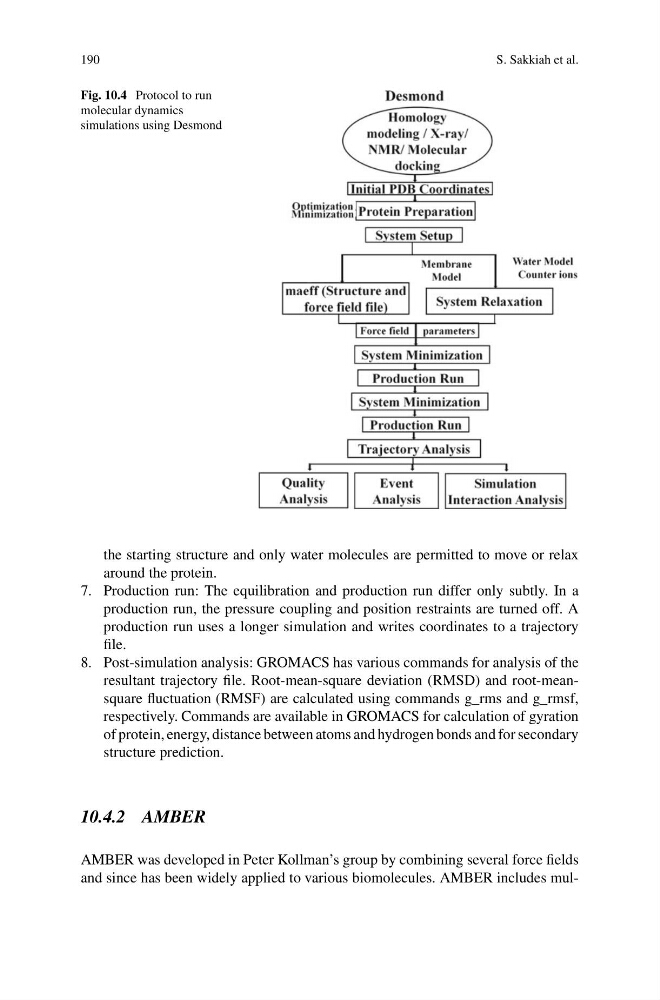
Fig. 10.4 Protocol to run
molecular dynamics
simulations using Desmond
the starting structure and only water molecules are permitted to move or relax
around the protein.
7. Production run: The equilibration and production run differ only subtly. In a
production run, the pressure coupling and position restraints are turned off. A
production run uses a longer simulation and writes coordinates to a trajectory
file.
8. Post-simulation analysis: GROMACS has various commands for analysis of the
resultant trajectory file. Root-mean-square deviation (RMSD) and root-meansquare fluctuation (RMSF) are calculated using commands g_rms and g_rmsf,
respectively. Commands are available in GROMACS for calculation of gyration
of protein, energy, distance between atoms and hydrogen bonds and for secondary
structure prediction.
10.4.2 AMBER
AMBER was developed in Peter Kollman’s group by combining several force fields
and since has been widely applied to various biomolecules. AMBER includes mul-