

52
this disorder then symptoms usually appear in early childhood. Children with this
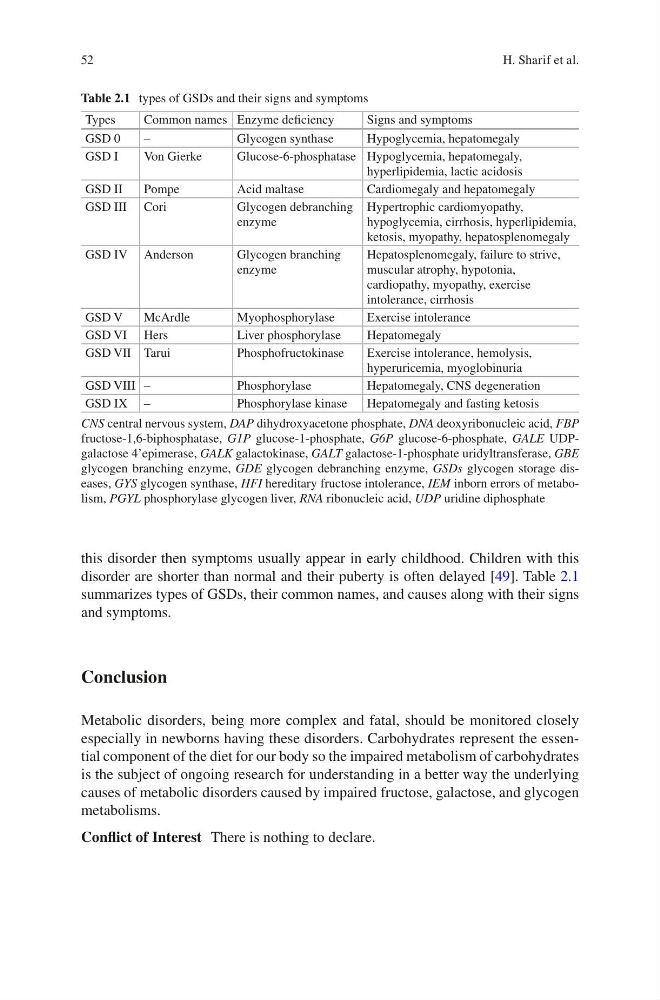
disorder are shorter than normal and their puberty is often delayed [49]. Table 2.1
summarizes types of GSDs, their common names, and causes along with their signs
and symptoms.
Conclusion
Metabolic disorders, being more complex and fatal, should be monitored closely
especially in newborns having these disorders. Carbohydrates represent the essential component of the diet for our body so the impaired metabolism of carbohydrates
is the subject of ongoing research for understanding in a better way the underlying
causes of metabolic disorders caused by impaired fructose, galactose, and glycogen
metabolisms.
Conflict of Interest There is nothing to declare.
Table 2.1 types of GSDs and their signs and symptoms
Types
Common names Enzyme deficiency
Signs and symptoms
GSD 0
–
Glycogen synthase
Hypoglycemia, hepatomegaly
GSD I
Von Gierke
Glucose-6-phosphatase Hypoglycemia, hepatomegaly,
hyperlipidemia, lactic acidosis
GSD II
Pompe
Acid maltase
Cardiomegaly and hepatomegaly
GSD III Cori
Glycogen debranching
enzyme
Hypertrophic cardiomyopathy,
hypoglycemia, cirrhosis, hyperlipidemia,
ketosis, myopathy, hepatosplenomegaly
GSD IV Anderson
Glycogen branching
enzyme
Hepatosplenomegaly, failure to strive,
muscular atrophy, hypotonia,
cardiopathy, myopathy, exercise
intolerance, cirrhosis
GSD V
McArdle
Myophosphorylase
Exercise intolerance
GSD VI Hers
Liver phosphorylase
Hepatomegaly
GSD VII Tarui
Phosphofructokinase
Exercise intolerance, hemolysis,
hyperuricemia, myoglobinuria
GSD VIII –
Phosphorylase
Hepatomegaly, CNS degeneration
GSD IX –
Phosphorylase kinase
Hepatomegaly and fasting ketosis
CNS central nervous system, DAP dihydroxyacetone phosphate, DNA deoxyribonucleic acid, FBP
fructose-1,6-biphosphatase, G1P glucose-1-phosphate, G6P glucose-6-phosphate, GALE UDPgalactose 4’epimerase, GALK galactokinase, GALT galactose-1-phosphate uridyltransferase, GBE
glycogen branching enzyme, GDE glycogen debranching enzyme, GSDs glycogen storage diseases, GYS glycogen synthase, HFI hereditary fructose intolerance, IEM inborn errors of metabolism, PGYL phosphorylase glycogen liver, RNA ribonucleic acid, UDP uridine diphosphate
H. Sharif et al.