

54
gmx_d solvate -cp newbox.gro -cs spc216.gro -o
solv.gro -p topol.top
In case a charged species is involved, placing of appropriate
counterions is needed in order to have a neutral net charge;
please refer to Note 4.
The solvated system should now be energy minimized before proceeding with the dynamics simulation. In order to do so, an energy
minimization protocol should be used. The energy minimization
protocol will be provided to the grompp module of gromacs suite
in order to produce a portable binary run input file of .tpr format.
The .tpr file contains information with regard to the starting structure of the simulation, as well as the molecular topology and all the
simulation parameters. The simulation parameters are described in
the protocol file em.mdp. An example protocol file (based on the
tutorials provided by Assist. Prof. Justin Lemkul at http://www.
mdtutorials.com/gmx/ but modified in order to be compatible
with the use of charmm36 force field (see Note 5)) is available in
the online version and presented in Fig. 6. The file contains selfexplanatory comments at each line.
1. The terminal command to be used in order to produce the
binary input file in accordance with the minimization protocol
is the following:
gmx_d grompp -f em.mdp -c solv.gro -p topol.
top -o em.tpr
In order to run the simulation, the mdrun module of the gromacs
suite should be used, which is its main computational chemistry
engine.
3.2 Energy
Minimization
3.2.1 Create Input
3.2.2 Run the Energy
Minimization Simulation
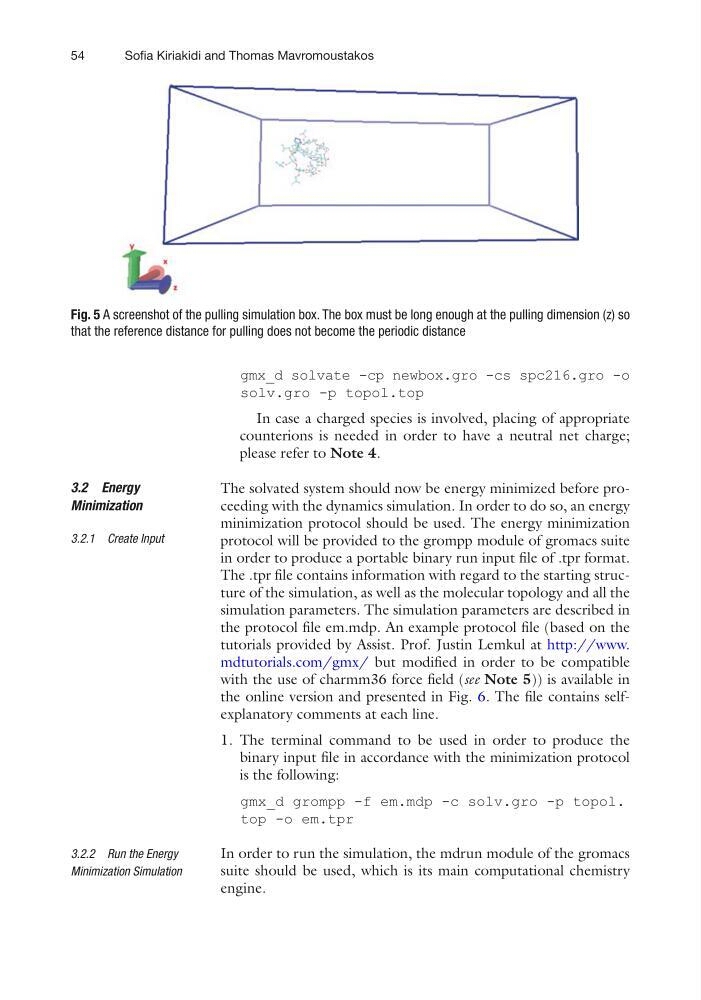
Fig. 5 A screenshot of the pulling simulation box. The box must be long enough at the pulling dimension (z) so
that the reference distance for pulling does not become the periodic distance
Sofia Kiriakidi and Thomas Mavromoustakos