

113
options that closely depend on the system under study. However,
based on personal experience, we indicate the following modifications for optimal performance.
6. Binding Site Bounding Box: Box sizes that range between 15
and 25Å (depending on the system’s size) are preferable.
Docking Engine: GADock may be preferred. The other options
are generally safe to be left as default.
7. In the same menu, click Advanced and increase Max. Generations
to 10,000. Press OK.
8. In the main Docking menu press Start to begin the docking
calculation.
9. After the calculation is over, you are provided with a docking
score (in kcal/mol) and a PDB structure of the complex in its
most favorable conformation (save it as complex.pdb).
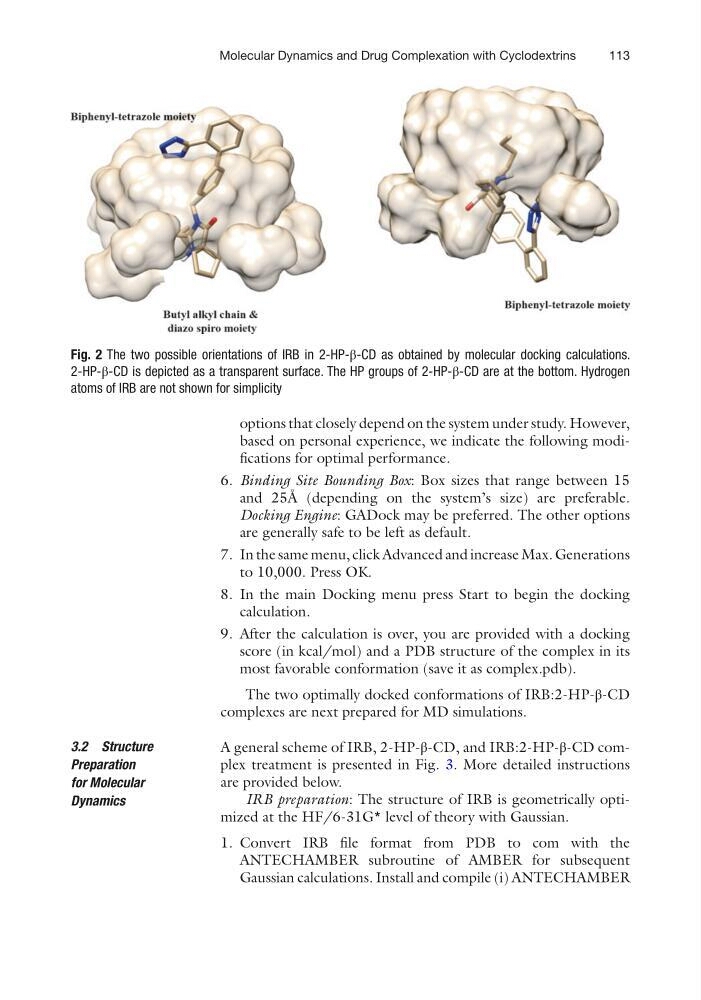
The two optimally docked conformations of IRB:2-HP-β-CD
complexes are next prepared for MD simulations.
A general scheme of IRB, 2-HP-β-CD, and IRB:2-HP-β-CD complex treatment is presented in Fig. 3. More detailed instructions
are provided below.
IRB preparation: The structure of IRB is geometrically optimized at the HF/6-31G* level of theory with Gaussian.
1. Convert IRB file format from PDB to com with the
ANTECHAMBER subroutine of AMBER for subsequent
Gaussian calculations. Install and compile (i) ANTECHAMBER
3.2 Structure
Preparation
for Molecular
Dynamics
Fig. 2 The two possible orientations of IRB in 2-HP-β-CD as obtained by molecular docking calculations.
2-HP-β-CD is depicted as a transparent surface. The HP groups of 2-HP-β-CD are at the bottom. Hydrogen
atoms of IRB are not shown for simplicity
Molecular Dynamics and Drug Complexation with Cyclodextrins