

260
9 Least-Squares Method
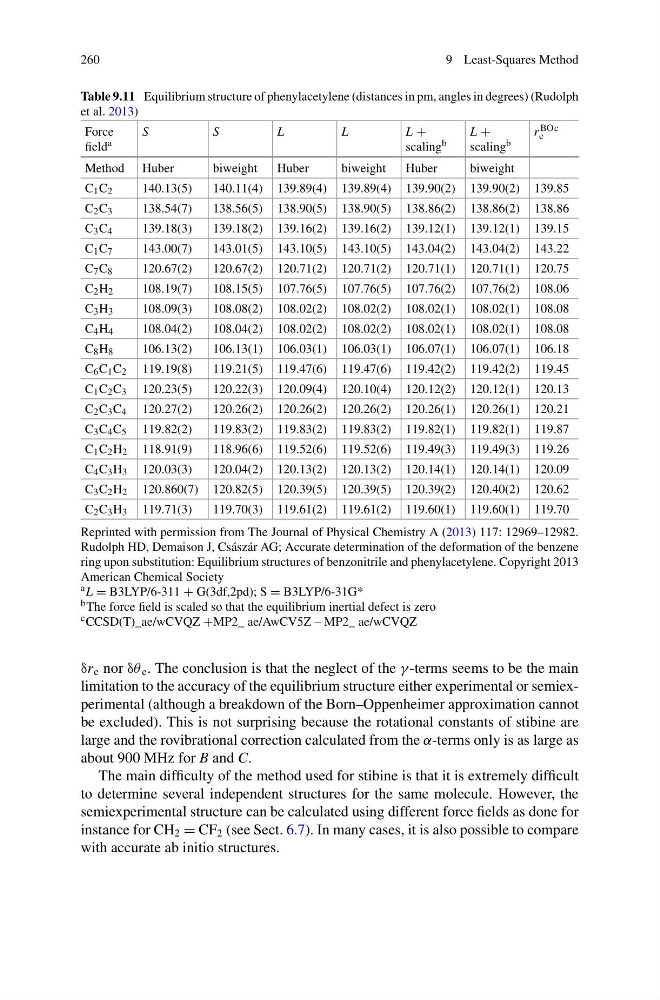
Table 9.11 Equilibrium structure of phenylacetylene (distances in pm, angles in degrees) (Rudolph
et al. 2013)
Force
field a
S
S
L
L
L +
scaling b
L +
scaling b
r BO
e
c
Method Huber
biweight
Huber
biweight
Huber
biweight
C 1 C 2
140.13(5)
140.11(4) 139.89(4) 139.89(4) 139.90(2) 139.90(2) 139.85
C 2 C 3
138.54(7)
138.56(5) 138.90(5) 138.90(5) 138.86(2) 138.86(2) 138.86
C 3 C 4
139.18(3)
139.18(2) 139.16(2) 139.16(2) 139.12(1) 139.12(1) 139.15
C 1 C 7
143.00(7)
143.01(5) 143.10(5) 143.10(5) 143.04(2) 143.04(2) 143.22
C 7 C 8
120.67(2)
120.67(2) 120.71(2) 120.71(2) 120.71(1) 120.71(1) 120.75
C 2 H 2
108.19(7)
108.15(5) 107.76(5) 107.76(5) 107.76(2) 107.76(2) 108.06
C 3 H 3
108.09(3)
108.08(2) 108.02(2) 108.02(2) 108.02(1) 108.02(1) 108.08
C 4 H 4
108.04(2)
108.04(2) 108.02(2) 108.02(2) 108.02(1) 108.02(1) 108.08
C 8 H 8
106.13(2)
106.13(1) 106.03(1) 106.03(1) 106.07(1) 106.07(1) 106.18
C 6 C 1 C 2 119.19(8)
119.21(5) 119.47(6) 119.47(6) 119.42(2) 119.42(2) 119.45
C 1 C 2 C 3 120.23(5)
120.22(3) 120.09(4) 120.10(4) 120.12(2) 120.12(1) 120.13
C 2 C 3 C 4 120.27(2)
120.26(2) 120.26(2) 120.26(2) 120.26(1) 120.26(1) 120.21
C 3 C 4 C 5 119.82(2)
119.83(2) 119.83(2) 119.83(2) 119.82(1) 119.82(1) 119.87
C 1 C 2 H 2 118.91(9)
118.96(6) 119.52(6) 119.52(6) 119.49(3) 119.49(3) 119.26
C 4 C 3 H 3 120.03(3)
120.04(2) 120.13(2) 120.13(2) 120.14(1) 120.14(1) 120.09
C 3 C 2 H 2 120.860(7) 120.82(5) 120.39(5) 120.39(5) 120.39(2) 120.40(2) 120.62
C 2 C 3 H 3 119.71(3)
119.70(3) 119.61(2) 119.61(2) 119.60(1) 119.60(1) 119.70
Reprinted with permission from The Journal of Physical Chemistry A (2013) 117: 12969–12982.
Rudolph HD, Demaison J, Császár AG; Accurate determination of the deformation of the benzene
ring upon substitution: Equilibrium structures of benzonitrile and phenylacetylene. Copyright 2013
American Chemical Society
a L = B3LYP/6-311 + G(3df,2pd); S = B3LYP/6-31G*
b The force field is scaled so that the equilibrium inertial defect is zero
c CCSD(T)_ae/wCVQZ +MP2_ ae/AwCV5Z – MP2_ ae/wCVQZ
δr e nor δθ e . The conclusion is that the neglect of the γ -terms seems to be the main
limitation to the accuracy of the equilibrium structure either experimental or semiexperimental (although a breakdown of the Born–Oppenheimer approximation cannot
be excluded). This is not surprising because the rotational constants of stibine are
large and the rovibrational correction calculated from the α-terms only is as large as
about 900 MHz for B and C.
The main difficulty of the method used for stibine is that it is extremely difficult
to determine several independent structures for the same molecule. However, the
semiexperimental structure can be calculated using different force fields as done for
instance for CH 2 = CF 2 (see Sect. 6.7). In many cases, it is also possible to compare
with accurate ab initio structures.