

alignment, there are 7 matches followed by a deletion from the reference, whereas the
deleted base is C and the last 5 bases are matches.
Example CIGAR: 3M1I3M1D5M
3M 3 matches/mismatches
1I
1 insertion
3M 3 matches/mismatches
1D 1 deletion
5M 5 matches/mismatches
Version 2 (new Version)
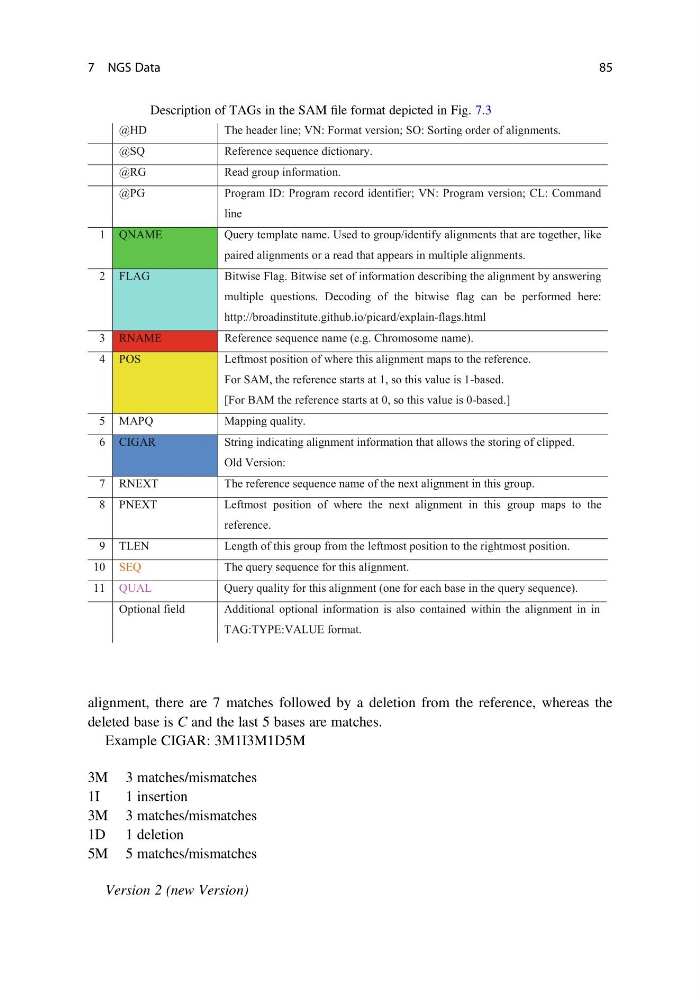
Table 7.2 Description of TAGs in the SAM file format depicted in Fig. 7.3
@HD
The header line; VN: Format version; SO: Sorting order of alignments.
@SQ
Reference sequence dictionary.
@RG
Read group information.
@PG
Program ID: Program record identifier; VN: Program version; CL: Command
line
QNAME
Query template name. Used to group/identify alignments that are together, like
paired alignments or a read that appears in multiple alignments.
FLAG
Bitwise Flag. Bitwise set of information describing the alignment by answering
multiple questions. Decoding of the bitwise flag can be performed here:
http://broadinstitute.github.io/picard/explain-flags.html
RNAME
Reference sequence name (e.g. Chromosome name).
POS
Leftmost position of where this alignment maps to the reference.
For SAM, the reference starts at 1, so this value is 1-based.
[For BAM the reference starts at 0, so this value is 0-based.]
MAPQ
Mapping quality.
CIGAR
String indicating alignment information that allows the storing of clipped.
Old Version:
RNEXT
The reference sequence name of the next alignment in this group.
PNEXT
Leftmost position of where the next alignment in this group maps to the
reference.
TLEN
Length of this group from the leftmost position to the rightmost position.
SEQ
The query sequence for this alignment.
1
2
3
4
5
6
7
8
9
10
11 QUAL
Query quality for this alignment (one for each base in the query sequence).
Optional field
Additional optional information is also contained within the alignment in in
TAG:TYPE:VALUE format.
7 NGS Data
85